standard-title
Drug Master File (DMF) & Regulatory Documentation Support
Taj Medico provides DMF preparation and pharmaceutical regulatory documentation services for global markets. We support API documentation, dossier compilation, regulatory submissions, stability data, and quality documentation.
Drug Master File (DMF) & Regulatory Documentation Support
Taj Medico provides DMF preparation and pharmaceutical regulatory documentation services for global markets. We support API documentation, dossier compilation, regulatory submissions, stability data, and quality documentation.
Drug Master File (DMF) and regulatory documentation are important components of pharmaceutical regulatory strategy, particularly for active pharmaceutical ingredients, excipients, manufacturing processes, and confidential technical information. A well-prepared regulatory package helps present accurate information to health authorities while protecting proprietary manufacturing and technical knowledge where applicable.
At Taj Medico, we support pharmaceutical businesses with DMF-related documentation and regulatory activities for pharmaceutical products and ingredients intended for domestic and international markets. Our approach focuses on technical accuracy, document consistency, regulatory organization, confidentiality, and appropriate lifecycle management.
DMF preparation requires coordination between manufacturing, quality control, quality assurance, analytical development, research and development, and regulatory affairs. The documentation must provide sufficient technical information to support regulatory assessment while remaining aligned with the actual manufacturing and quality-control processes.
What Is a Drug Master File (DMF)?
A Drug Master File is a regulatory submission containing detailed information about facilities, processes, materials, or other information related to the manufacture, processing, packaging, and storage of a pharmaceutical component or product.
The exact definition, format, submission process, and regulatory expectations vary between jurisdictions.
A DMF may contain confidential information that a manufacturer or supplier does not wish to disclose directly to a customer. Regulatory authorities can review the confidential information while the customer receives appropriate authorization or supporting documentation.
Depending on the regulatory framework and product, master-file concepts may be used for information concerning:
Active pharmaceutical ingredients
Manufacturing processes
Excipients
Packaging components
Facilities
Other confidential technical information
Importance of DMF Documentation in Pharmaceutical Registration
DMF documentation can help pharmaceutical companies demonstrate appropriate control over critical materials and manufacturing processes.
A well-organized DMF can support:
Regulatory submissions
API registration
Product registration
Confidential manufacturing information
Quality assessment
Supplier qualification
Regulatory review
Product lifecycle management
The specific purpose of a DMF depends on the applicable regulatory system and the type of information being submitted.
DMF Preparation Services
Preparing a DMF requires the systematic collection and organization of technical information from multiple departments.
Our DMF-support activities may include:
Technical-document collection
Document gap assessment
Regulatory document review
Manufacturing information compilation
Analytical information compilation
Specification review
Stability-data organization
Quality documentation
Regulatory formatting
Submission-support documentation
Lifecycle updates
We focus on maintaining consistency between the DMF and the underlying manufacturing and quality systems.
Types of Drug Master Files
DMF classifications vary between regulatory jurisdictions. The terminology and submission categories used by one authority may not be identical to those used by another.
Common areas covered by master-file systems may include:
API Master File
An API master file provides detailed information about the manufacture and control of an active pharmaceutical ingredient.
It can include:
General information
Manufacturing process
Process controls
Starting materials
Impurity profile
Process validation
Specifications
Analytical procedures
Reference standards
Packaging
Stability
Excipient Documentation
Certain regulatory systems may allow or require detailed confidential information regarding excipients or specialized pharmaceutical ingredients.
Documentation can address:
Manufacturing process
Quality controls
Specifications
Analytical procedures
Stability
Packaging
Packaging-Related Master Documentation
For specific regulatory frameworks, confidential information concerning packaging components may require dedicated technical documentation.
The exact requirements depend on the material and regulatory authority.
DMF Content and Technical Information
A DMF should contain information appropriate to its purpose and regulatory pathway.
Technical sections can include:
General Information
This section may identify:
Substance or material
Manufacturer
Manufacturing location
Product characteristics
Regulatory purpose
Manufacturing Process
The manufacturing section can describe:
Manufacturing steps
Process controls
Starting materials
Critical process parameters
Process flow
In-process controls
Process validation information
Quality Control
Quality information can cover:
Specifications
Analytical methods
Method validation
Impurity controls
Reference standards
Batch analysis
Packaging and Storage
The documentation can include information concerning:
Container-closure systems
Packaging materials
Storage conditions
Transportation considerations
Stability
The exact information required should be determined according to the applicable regulatory framework.
API Regulatory Documentation
Active pharmaceutical ingredients are a major focus of regulatory documentation in pharmaceutical manufacturing.
API documentation can provide information about:
Chemical identity
Structure
Manufacturing process
Starting materials
Raw materials
Process controls
Impurity profile
Specifications
Analytical procedures
Batch analysis
Stability
Packaging
Accurate API documentation helps regulators assess whether the active ingredient is appropriately manufactured and controlled.
API Manufacturing Process Documentation
The manufacturing process section should accurately represent how the API is manufactured.
Information may include:
Starting materials
Manufacturing steps
Reaction conditions
Process controls
Isolation and purification
Drying
Milling where applicable
Packaging
Storage
Critical manufacturing parameters should be identified and controlled according to the process and applicable quality requirements.
API Impurity and Control Strategy
Impurity control is an important component of pharmaceutical regulatory documentation.
Impurities can arise from:
Starting materials
Manufacturing reactions
Process intermediates
Side reactions
Degradation
Residual solvents
Reagents and catalysts
The DMF should provide an appropriate explanation of the impurity profile and the controls used to maintain impurity levels within established limits.
Analytical Method Documentation
Analytical procedures provide the means to evaluate the quality of pharmaceutical substances and products.
DMF documentation may contain methods for:
Identification
Assay
Related substances
Residual solvents
Water content
Elemental impurities where applicable
Other product-specific tests
Analytical methods should be appropriately developed, validated, verified, or otherwise demonstrated to be suitable for their intended purpose according to applicable requirements.
Analytical Method Validation Documentation
Where required, validation documentation may address characteristics such as:
Specificity
Accuracy
Precision
Linearity
Range
Robustness
Detection limit
Quantitation limit
The validation strategy should correspond to the intended analytical use and relevant regulatory expectations.
DMF Stability Documentation
Stability information is an important component of many pharmaceutical regulatory packages.
Stability studies can provide information about how an API or other pharmaceutical material behaves under defined storage conditions.
Relevant parameters may include:
Assay
Related substances
Appearance
Water content
Physical characteristics
Other product-specific quality attributes
Stability information can support proposed storage conditions and retest periods where applicable.
DMF Gap Assessment
A DMF gap assessment identifies missing, outdated, inconsistent, or insufficient documentation before regulatory submission.
We can review areas such as:
Manufacturing process
Specifications
Analytical methods
Validation
Impurity information
Stability
Packaging
Batch data
Certificates
Administrative documentation
Early gap identification can help reduce avoidable delays during regulatory review.
Regulatory Dossier and DMF Integration
A DMF may be referenced by a pharmaceutical product dossier rather than duplicating confidential information within the applicant’s submission.
The product dossier and DMF should remain technically consistent.
Areas that may require alignment include:
API specifications
Manufacturer information
Manufacturing process
Analytical methods
Impurity controls
Stability information
Packaging
Storage conditions
Inconsistencies between regulatory documents can generate questions during assessment, making careful cross-document review important.
Letter of Authorization and Regulatory Access
Certain regulatory systems use authorization mechanisms that allow a regulatory authority to access confidential master-file information in support of another application.
The specific terminology and procedure vary by jurisdiction.
Regulatory documentation may therefore need to include appropriate authorization information when a DMF is referenced by a pharmaceutical applicant.
DMF Regulatory Submission Support
Regulatory submission requirements vary by health authority.
Our submission-support activities may include:
Document compilation
Regulatory formatting
Technical review
Administrative-document coordination
Submission readiness assessment
Cross-reference checking
Regulatory correspondence
Deficiency-response support
Lifecycle updates
We adapt documentation to the applicable regulatory framework rather than applying a single universal format.
CTD and DMF Documentation
The Common Technical Document (CTD) is widely used for pharmaceutical regulatory submissions, while master-file systems have their own requirements depending on the jurisdiction.
Technical information from a DMF may support relevant sections of a pharmaceutical dossier, particularly information related to the drug substance.
We coordinate information so that the DMF and product dossier remain consistent where cross-referencing is required.
Regulatory Deficiency Response Support
Regulatory authorities may raise questions regarding manufacturing processes, specifications, analytical methods, impurities, stability, or other technical areas.
We support structured responses by coordinating information from relevant technical and quality teams.
A deficiency response may require:
Technical clarification
Additional analytical data
Revised specifications
Manufacturing information
Validation documentation
Stability results
Updated regulatory sections
Responses should be complete, scientifically justified, and consistent with the submitted documentation.
DMF Lifecycle Management
A DMF is not necessarily a static document. Changes to manufacturing processes, specifications, facilities, analytical methods, packaging, or other technical information may require updates.
Lifecycle activities can include:
DMF amendments
Technical updates
Manufacturing-site changes
Process changes
Specification changes
Analytical-method changes
Packaging changes
Stability updates
Regulatory correspondence
Each change should be assessed according to the relevant regulatory requirements.
DMF Change Control
Changes affecting master-file information should be managed through an appropriate quality and regulatory change-control system.
The impact assessment may consider:
Product quality
Regulatory commitments
Stability
Analytical testing
Manufacturing
Validation
Referenced product dossiers
The applicable regulatory notification or approval pathway depends on the nature and significance of the change.
Confidentiality in DMF Management
Confidentiality is particularly important for master-file information because the documentation may contain proprietary manufacturing and technical information.
A controlled documentation system can help protect:
Manufacturing processes
Process parameters
Proprietary formulations
Analytical information
Supplier information
Technical specifications
Intellectual property
Access to confidential documentation should be appropriately controlled according to organizational procedures and regulatory requirements.
DMF Documentation for Global Pharmaceutical Markets
Pharmaceutical companies operating internationally may need master-file or equivalent technical documentation for different regulatory authorities.
Requirements can differ regarding:
Document format
Submission procedures
Technical content
Administrative information
Authorization mechanisms
Updates
Regulatory correspondence
Lifecycle management
We support international regulatory documentation by reviewing the requirements applicable to the target market.
DMF Support for Pharmaceutical Manufacturers and Suppliers
API manufacturers, pharmaceutical companies, contract manufacturers, and suppliers may require regulatory documentation support at different stages of the product lifecycle.
Our services can support:
New DMF preparation
Existing DMF updates
Regulatory gap assessment
API documentation
Analytical documentation
Stability documentation
Manufacturing information
Regulatory submission support
Deficiency responses
Lifecycle management
Quality Review of DMF Documentation
Before submission or regulatory use, a detailed quality review helps identify inconsistencies and missing information.
The review may examine:
Product identity
Manufacturing-site information
Specifications
Analytical methods
Batch-analysis data
Impurity information
Stability results
Packaging details
Cross-references
Document versions
A thorough review improves the overall quality and consistency of the regulatory package.
Why Choose Taj Medico for DMF & Regulatory Documentation Support?
We provide DMF and pharmaceutical regulatory documentation support focused on technical accuracy, structured document management, quality, and regulatory consistency.
Our capabilities include:
DMF preparation support
API documentation
Regulatory gap assessment
Quality documentation
Manufacturing-process documentation
Analytical-method documentation
Stability documentation
Regulatory submission support
Deficiency-response coordination
DMF lifecycle management
Change-control documentation
Regulatory document review
We coordinate technical information from manufacturing, quality control, quality assurance, analytical, and regulatory functions to help establish a complete documentation package.
Supporting International Pharmaceutical Registration
For pharmaceutical businesses expanding into international markets, regulatory documentation can become increasingly complex. Each destination market may have specific requirements for APIs, finished products, manufacturing facilities, certificates, dossiers, and supporting technical information.
We support businesses by organizing regulatory documentation around the applicable requirements and ensuring that relevant technical information remains consistent across connected submissions.
Drug Master File (DMF) and regulatory documentation support are important for pharmaceutical companies seeking to demonstrate appropriate control over active pharmaceutical ingredients, manufacturing processes, analytical procedures, and other confidential technical information.
A well-prepared master file requires accurate manufacturing information, appropriate specifications, analytical documentation, impurity controls, stability data, and regulatory organization. It must also remain consistent with associated pharmaceutical dossiers and manufacturing practices.
At Taj Medico, we provide structured DMF preparation and regulatory documentation support for pharmaceutical businesses and global buyers. Our services cover technical-document compilation, gap assessment, API documentation, analytical information, stability data, regulatory submission support, deficiency responses, and lifecycle management.
By maintaining accurate, controlled, and well-organized regulatory documentation, pharmaceutical companies can establish a stronger foundation for product registration, regulatory assessment, manufacturing compliance, and international market expansion.
Colistimethate Sodium for Injection IP 3 MIU Manufacturer & Supplier in India Colistimethate Sodium for Injection IP 3 MIU Manufacturer in India We, at Taj Medico, are engaged in the manufacturing and supply of pharmaceutical products for domestic and international markets. Our Colistimethate Sodium for Injection IP 3 MIU is manufactured with a strong focus on quality, consistency, appropriate packaging, and reliable supply. Colistimethate sodium is a prodrug of colistin and belongs to the polymyxin class of antibiotics. It is used in clinical settings for the management of certain serious bacterial infections caused by susceptible organisms. As a Colistimethate Sodium for Injection IP 3 MIU Manufacturer in India, we understand the importance of maintaining consistent manufacturing standards for injectable pharmaceutical products. Our approach combines controlled production processes, quality checks, suitable packaging, and organized supply-chain management to support healthcare and pharmaceutical partners. For buyers searching for a Colistimethate Injection Manufacturer in Mumbai, Colistimethate Sodium Manufacturer in Vapi, Gujarat, or a dependable PAN India supplier and exporter, Taj Medico provides a comprehensive pharmaceutical supply solution. What Is Colistimethate Sodium for Injection IP 3 MIU? Colistimethate Sodium for Injection IP 3 MIU contains colistimethate sodium, which is converted in the body to the […]
Colistimethate Sodium for Injection IP 4.5 MIU Manufacturer & Supplier in India – Taj Medico We at Taj Medico are engaged in the manufacturing and supply of pharmaceutical products, including Colistimethate Sodium for Injection IP 4.5 MIU. Based in Vapi, Gujarat, India, we support pharmaceutical distributors, institutional buyers, healthcare businesses, and authorized international customers seeking dependable pharmaceutical sourcing. Colistimethate sodium is a polymyxin antibiotic used in selected serious infections caused by susceptible Gram-negative bacteria. Because it is generally reserved for difficult-to-treat infections, appropriate product quality, professional medical supervision, and responsible antibiotic use are important considerations. Our focus is to provide pharmaceutical products through controlled manufacturing processes, quality-oriented practices, suitable packaging, and organized domestic and international supply. Colistimethate Sodium for Injection IP 4.5 MIU Manufacturer in India Colistimethat Sodium for Injection IP 4.5 MIU As a Colistimethate Sodium for Injection IP 4.5 MIU manufacturer in India, we understand the importance of consistency in injectable pharmaceutical manufacturing. Injectable medicines require controlled production environments, appropriate handling procedures, accurate filling, quality testing, and suitable packaging. Colistimethate sodium is a prodrug that is converted to colistin in the body. Colistin belongs to the polymyxin class of antibiotics and has activity against certain Gram-negative organisms. It […]
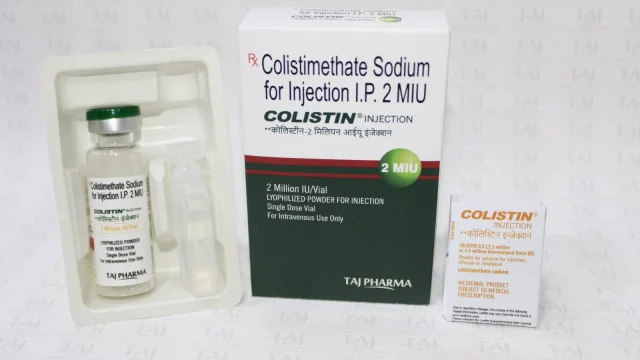
Colistimethate Sodium for Injection IP 2MIU Manufacturer & Supplier in India Taj Medico is an established Colistimethate Sodium for Injection IP 2MIU manufacturer, supplier, and exporter in India, serving authorized pharmaceutical businesses and healthcare procurement channels from Vapi, Gujarat. We focus on quality-oriented pharmaceutical manufacturing, controlled processes, appropriate packaging, documentation, and dependable supply coordination for domestic and eligible international markets. Colistimethate Sodium for Injection IP 2MIU is an injectable antibacterial formulation containing colistimethate sodium, a prodrug of colistin. It is used under qualified medical supervision for selected serious infections caused by susceptible Gram-negative bacteria. Its clinical use is generally associated with hospital and specialist-care settings, particularly where antimicrobial resistance can limit available treatment choices. 🧪 Colistimethate Sodium for Injection IP 2MIU – Product Overview Colistimethate Sodium for Injection IP 2MIU Colistimethate Sodium belongs to the polymyxin class of antibacterial medicines and serves as a prodrug of colistin. Following administration, it is converted to active colistin, which exerts antibacterial activity against susceptible organisms. The 2MIU strength represents two million international units. The appropriate dosage and administration schedule are determined by qualified healthcare professionals based on the patient’s clinical condition, infection characteristics, kidney function, microbiological information, and other relevant factors. As a […]
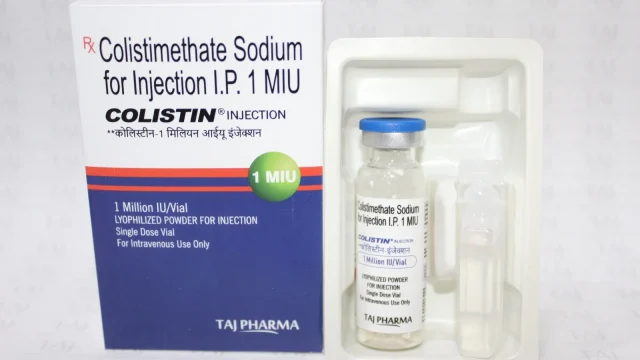
Colistimethate Sodium for Injection IP 1MIU Manufacturer & Supplier in India Taj Medico is a well-established pharmaceutical manufacturer, exporter, and supplier of Colistimethate Sodium for Injection IP 1MIU in India, serving pharmaceutical businesses and authorized healthcare supply channels from Vapi, Gujarat, India. With an emphasis on quality-oriented manufacturing, controlled processes, product consistency, and dependable supply support, we provide pharmaceutical solutions for domestic and eligible international markets. Colistimethate Sodium for Injection IP 1MIU, also known as a colistimethate or colistin injectable formulation, is an important antibacterial medicine used in the management of certain serious infections caused by susceptible Gram-negative bacteria. It is generally used in hospital settings under the supervision of qualified healthcare professionals, particularly when infections involve organisms with limited treatment options. 🧪 Colistimethate Sodium for Injection IP 1MIU – Product Overview Colistimethate Sodium for Injection Colistimethate Sodium is a prodrug of colistin and belongs to the polymyxin group of antibacterial medicines. The injectable formulation is used when systemic antibacterial treatment is medically indicated for appropriate susceptible infections. The 1 MIU strength represents one million international units and is supplied for professional medical use according to approved product information and applicable treatment protocols. The dosage, frequency, duration, and method of […]
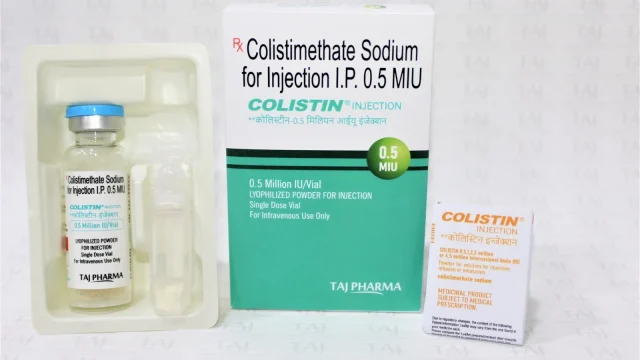
Colistimethate Sodium for Injection IP 0.5MIU Manufacturer Supplier in India – Why Taj Medico is the Best Exporter for Global Buyers Colistimethate Sodium for Injection IP 0.5MIU Manufacturer in India We are a trusted pharmaceutical manufacturer, exporter, and supplier of Colistimethate Sodium for Injection IP 0.5MIU from Vapi, Gujarat, India. Our focus is on supplying quality-assured finished pharmaceutical formulations that meet the expectations of hospitals, healthcare distributors, pharmaceutical companies, and international buyers. Colistimethate Sodium is a polymyxin-class antibacterial medicine used primarily in serious infections caused by susceptible Gram-negative bacteria, particularly when resistance limits the usefulness of other antibiotics. Because colistimethate is generally reserved for severe infections, consistent pharmaceutical quality, appropriate manufacturing controls, accurate strength, and reliable supply are especially important. We manufacture and supply Colistimethate 0.5MIU (Colistin) Injection for domestic and international pharmaceutical markets, supporting buyers looking for dependable Indian manufacturing capabilities. Our approach combines controlled production, quality management, suitable packaging, documentation support, and professional export coordination. Colistimethate 0.5MIU (Colistin) Injection – Product Overview Colistimethate Sodium for Injection IP 0.5MIU Colistimethate Sodium for Injection IP 0.5MIU contains colistimethate sodium, a prodrug of colistin. After administration, colistimethate is converted in the body to active colistin, which acts against susceptible Gram-negative organisms […]
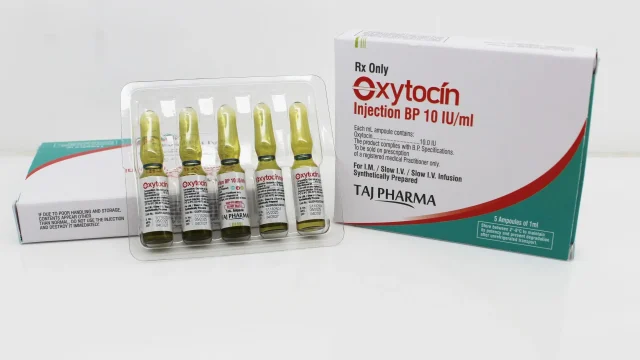
Oxytocin Injection 10 IU/ml Manufacturer and Supplier in India – Taj Medico for Global Pharmaceutical Buyers Taj Medico is a well-established Oxytocin Injection 10 IU/ml manufacturer, supplier, and exporter in India, serving pharmaceutical distributors, healthcare procurement organizations, hospitals, and international buyers. We focus on delivering pharmaceutical products manufactured and handled according to appropriate quality and regulatory requirements, with particular attention to product consistency, packaging, documentation, and supply-chain reliability. Oxytocin Injection 10 IU/ml is a prescription medicine used in obstetric care under professional medical supervision. Oxytocin is a synthetic form of a naturally occurring hormone that promotes uterine contractions. Depending on the clinical situation, healthcare professionals may use it to induce or augment labor and to help manage postpartum bleeding associated with inadequate uterine contraction. Because oxytocin can produce powerful uterine contractions, its administration requires appropriate medical assessment, dosing, monitoring, and supervision. Oxytocin Injection 10 IU/ml Manufacturer in India Oxytocin Injection 10 IU/ml As an Oxytocin Injection 10 IU/ml manufacturer in India, we understand the importance of maintaining reliable standards throughout pharmaceutical production and supply. Injectable medicines require particular attention to sterility, manufacturing controls, container integrity, labeling, storage, and transportation. We work to provide pharmaceutical buyers with a dependable supply of […]
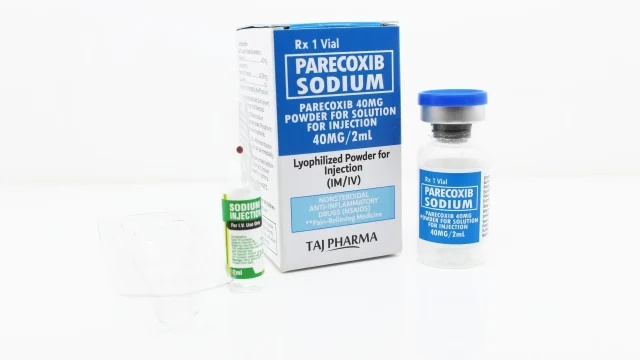
Parecoxib Injection 40mg Manufacturer and Supplier in India Parecoxib Injection 40mg is a prescription injectable analgesic belonging to the selective cyclooxygenase-2 (COX-2) inhibitor class. It is used in clinical settings for the short-term management of acute postoperative pain when parenteral treatment is appropriate. We provide pharmaceutical sourcing and supply solutions for authorized healthcare institutions, distributors, wholesalers, pharmaceutical companies, and international buyers seeking a dependable Parecoxib Injection 40mg manufacturer and supplier in India. Taj Medico operates from Vapi, Gujarat, India, a region with an established pharmaceutical manufacturing and industrial ecosystem. Our focus is on quality-oriented pharmaceutical production, responsible supply, appropriate documentation, and professional customer support for domestic and international markets. Parecoxib Injection 40mg Manufacturer in India Parecoxib Injection 40mg We supply Parecoxib Injection 40mg for eligible pharmaceutical and healthcare buyers requiring a reliable source from India. Injectable medicines demand careful attention to manufacturing controls, product specifications, packaging integrity, quality testing, and supply-chain management. Our pharmaceutical supply process is structured around these requirements. We work to maintain consistent product quality and support customers with relevant product and commercial information according to their market requirements. Parecoxib is a prodrug that is converted in the body to valdecoxib, its active component. As a selective […]
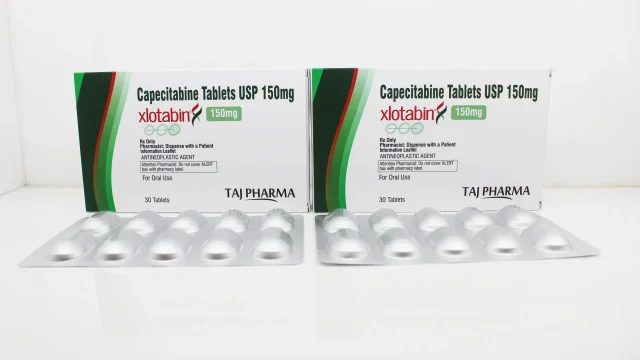
Capecitabine Tablets 150mg Manufacturer Supplier in India Capecitabine Tablets 150mg are an important oral chemotherapy medicine used in the management of several types of cancer. We at Taj Medico focus on supplying pharmaceutical products with consistent manufacturing standards, reliable quality, and professional support for domestic and international buyers. As a Capecitabine Tablets 150mg Manufacturer and Supplier in India, we serve pharmaceutical distributors, institutional buyers, healthcare procurement organizations, and eligible business partners across India and international markets. Capecitabine is an oral antimetabolite medicine that is converted in the body into 5-fluorouracil (5-FU), an active anticancer substance. Depending on the treatment protocol, it may be prescribed for cancers such as breast cancer, colorectal cancer, colon cancer, and rectal cancer, either alone or alongside other anticancer medicines. Treatment decisions, dosage, and monitoring should always be determined by a qualified oncology professional. Capecitabine Tablets 150mg Manufacturer in India Capecitabine Tablets USP 150mg We manufacture and supply Capecitabine Tablets 150mg with attention to product quality, manufacturing consistency, packaging, and supply-chain requirements. Our operations are based in Vapi, Gujarat, India, an established pharmaceutical manufacturing hub. For pharmaceutical buyers searching for a Capecitabine 150mg manufacturer in India, choosing an experienced supplier is important because oncology medicines require […]
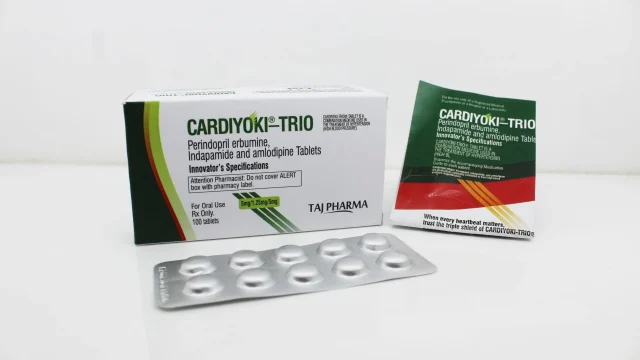
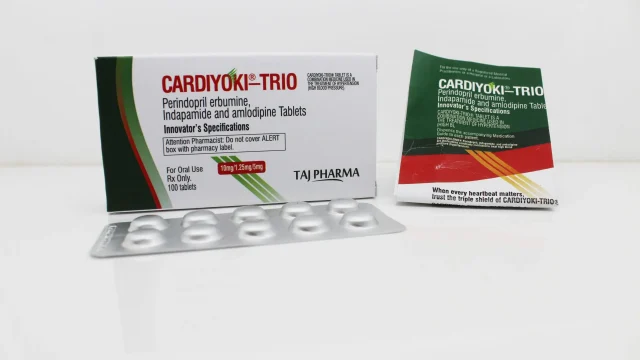
Perindopril Erbumine 5mg, Indapamide 1.25mg and Amlodipine 5mg Tablets Manufacturer & Supplier in India Perindopril Erbumine 5mg Indapamide 1.25mg Amlodipine 5mg Tablets Manufacturer in India We are an established manufacturer, supplier, and exporter of Perindopril Erbumine 5mg, Indapamide 1.25mg and Amlodipine 5mg Tablets in India, serving pharmaceutical distributors, healthcare businesses, institutional buyers, and international markets. Our focus is on maintaining consistent product quality, reliable manufacturing practices, appropriate packaging, and dependable supply support for buyers seeking cardiovascular and antihypertensive medicines. This triple-combination formulation brings together three established antihypertensive medicines: Perindopril Erbumine, an ACE inhibitor; Indapamide, a diuretic with antihypertensive properties; and Amlodipine, a calcium channel blocker. The combination is designed for the management of hypertension, particularly where treatment with multiple mechanisms may be clinically appropriate. We manufacture and supply products under applicable pharmaceutical quality requirements and support domestic as well as export-oriented requirements from our operations in Vapi, Gujarat, India. Cardiyoki-Trio Tablets – Triple-Combination Antihypertensive Medicine Perindopril erbumine 5mg Indapamide 1.25mg Amlodipine 5mg Tablets Perindopril Erbumine 5mg + Indapamide 1.25mg + Amlodipine 5mg Tablets, also referred to as Cardiyoki-Trio in the supplied product description, combine three medicines that lower blood pressure through complementary mechanisms. Perindopril Erbumine belongs to the angiotensin-converting enzyme […]
Perindopril Erbumine 10mg, Indapamide 1.25mg, Amlodipine 5mg Tablets Manufacturer & Supplier in India Taj Medico is an established pharmaceutical manufacturer, supplier, and exporter from Vapi, Gujarat, India, offering a broad range of finished pharmaceutical formulations for domestic and international markets. Among its cardiovascular product portfolio is Perindopril Erbumine 10mg, Indapamide 1.25mg, Amlodipine 5mg Tablets (Cardiyoki-Trio), a prescription fixed-dose combination formulated around three established antihypertensive medicines. The combination brings together perindopril erbumine, indapamide, and amlodipine, medicines that lower blood pressure through complementary pharmacological mechanisms. Perindopril belongs to the angiotensin-converting enzyme (ACE) inhibitor class, indapamide is a thiazide-like diuretic, and amlodipine is a calcium-channel blocker. Combining different antihypertensive mechanisms can provide a comprehensive approach to blood-pressure management when a multi-drug regimen is clinically appropriate. The exact approved indication, strength, formulation, and prescribing information should always be confirmed against the current product registration and approved dossier. For pharmaceutical distributors, institutional buyers, healthcare procurement organizations, and PCD pharma franchise partners, we focus on dependable manufacturing, consistent product quality, appropriate packaging, documentation support, and supply coordination. Perindopril Erbumine 10mg, Indapamide 1.25mg, Amlodipine 5mg Tablets Manufacturer in India Perindopril Erbumine , Indapamide, Amlodipine Tablets As a Perindopril Erbumine 10mg, Indapamide 1.25mg, Amlodipine 5mg Tablets Manufacturer in […]
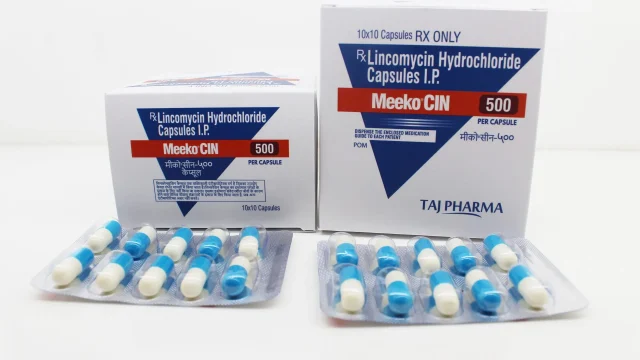
Lincomycin 500mg Capsule Manufacturer & Supplier in India Lincomycin 500mg Capsule Manufacturer in India Taj Medico is an established Lincomycin 500mg Capsule Manufacturer, Supplier, and Exporter in India, supporting authorized pharmaceutical distributors, healthcare businesses, institutional purchasers, and eligible international buyers. We focus on quality-oriented pharmaceutical manufacturing, consistent product standards, dependable supply, and professional customer service. Our manufacturing and supply operations are based in Vapi, Gujarat, an established pharmaceutical and industrial hub. From this location, we support customers looking for a Lincomycin 500mg Capsule Manufacturer in Vapi, Gujarat, while our distribution capabilities enable supply to customers across India. For businesses searching for Lincomycin 500mg Capsule Manufacturers PAN India, Lincomycin 500mg Capsule Suppliers in India, or a reliable pharmaceutical exporter, Taj Medico provides a structured procurement and supply channel. Lincomycin 500mg Capsule – Pharmaceutical Overview Lincomycin 500mg capsule Lincomycin is an antibiotic belonging to the lincosamide class. It works by inhibiting bacterial protein synthesis, thereby preventing susceptible bacteria from growing and multiplying. It is generally reserved for infections caused by susceptible bacteria when its use is considered medically appropriate. Depending on the approved product information and clinical circumstances, lincomycin may be used for certain serious bacterial infections involving susceptible organisms. The appropriate […]
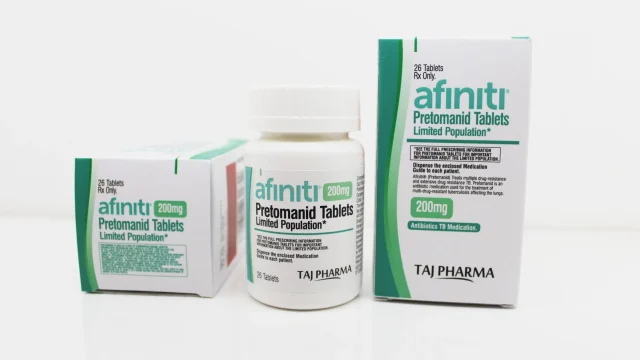
Pretomanid 200mg Tablets Manufacturer Supplier in India – Taj Medico Pretomanid 200mg Tablets are a specialized prescription antimycobacterial medicine used as part of combination therapy for certain forms of drug-resistant pulmonary tuberculosis (TB). Pretomanid belongs to the nitroimidazooxazine class and is not intended to be used as a standalone treatment for tuberculosis. According to current U.S. prescribing information, pretomanid 200mg is used together with bedaquiline and linezolid for a defined population of adults with drug-resistant pulmonary TB. Taj Medico is an established pharmaceutical manufacturer, supplier, and exporter based in Vapi, Gujarat, India, supporting qualified pharmaceutical businesses, distributors, healthcare institutions, and international buyers. We provide organized pharmaceutical sourcing solutions covering bulk requirements, domestic distribution, potential PCD Pharma Franchise opportunities, and export inquiries, subject to applicable regulatory requirements. Our objective is to provide dependable product coordination, appropriate packaging, professional customer communication, and supply support for customers seeking Pretomanid 200mg Tablets from India. Pretomanid 200mg Tablets Manufacturer in India Pretomanid 200mg Tablets As an Indian pharmaceutical company, we understand the importance of dependable sourcing for specialized medicines. Pretomanid 200mg Tablets Manufacturer in India is a high-intent pharmaceutical search term, but responsible product sourcing requires more than simply comparing supplier names or prices. Buyers […]
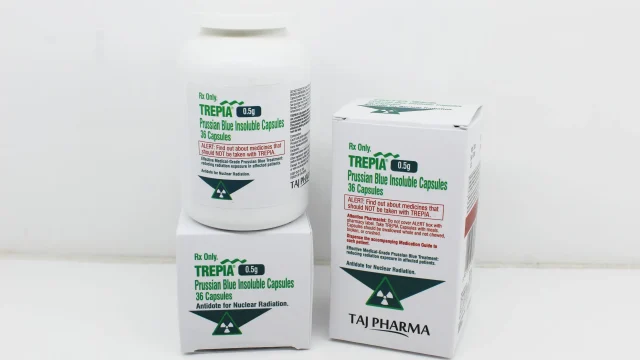
Prussian Blue Insoluble Capsules 0.5g Manufacturer, Supplier & PCD Pharma Franchise in India Prussian Blue Insoluble Capsules 0.5g are a specialized pharmaceutical formulation used under medical supervision for the management of certain cases of radioactive cesium or thallium contamination and non-radioactive thallium poisoning. Because this is a specialized medicine, dependable pharmaceutical manufacturing, controlled quality standards, appropriate packaging, and a reliable supply chain are important considerations for healthcare institutions, distributors, and pharmaceutical businesses. Taj Medico is an established manufacturer, exporter, and supplier of Prussian Blue Insoluble Capsules 0.5g from Vapi, Gujarat, India. We focus on supplying pharmaceutical products through professional distribution channels while supporting domestic and international requirements. Our approach emphasizes quality-oriented manufacturing, consistent product handling, dependable business communication, and efficient pharmaceutical distribution. Prussian Blue Insoluble Capsules 0.5g Manufacturer in India Prussian Blue Insoluble Capsules 0.5g We provide pharmaceutical sourcing support for customers looking for Prussian Blue Insoluble Capsules 0.5g manufacturers and suppliers in India. Specialized medicines require particular attention to product specifications, manufacturing controls, packaging, storage, documentation, and distribution. Prussian Blue Insoluble is not an ordinary pharmaceutical product intended for routine self-treatment. It is used for specific medical circumstances and should be administered according to appropriate professional guidance. We therefore […]
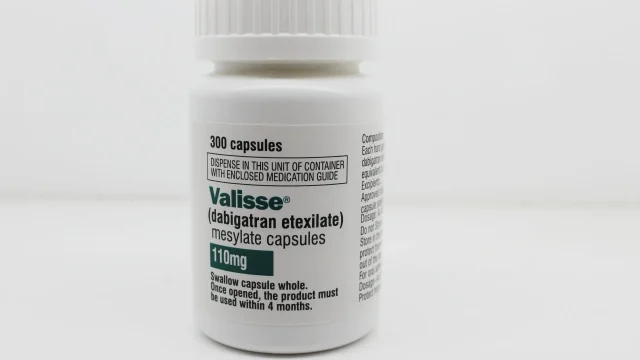
Dabigatran Etexilate Mesylate Capsules 110mg Manufacturer, Supplier and Exporter in India Dabigatran Etexilate Mesylate Capsules 110mg are prescription oral anticoagulant capsules used in approved indications for preventing and treating blood clots. We provide pharmaceutical sourcing solutions for businesses seeking dependable Dabigatran Capsules 110mg manufacturers, suppliers, exporters, and PCD Pharma Franchise opportunities in India. Our focus is on maintaining reliable pharmaceutical supply, appropriate product documentation, professional customer support, and market-oriented distribution. For buyers searching for Dabigatran Etexilate Mesylate Capsules 110mg manufacturers in India, we offer a structured approach to domestic and international pharmaceutical procurement. Dabigatran etexilate is converted in the body to dabigatran, an active direct thrombin inhibitor. By inhibiting thrombin, dabigatran interferes with the blood-clotting process and helps reduce the formation of certain blood clots. The 110mg strength is used in specific approved clinical situations according to the applicable prescribing information and regulatory requirements. Dabigatran Etexilate Mesylate Capsules 110mg Manufacturer in India Dabigatran Etexilate Mesylate Capsules 110mg (Valisse) We serve pharmaceutical buyers looking for a reliable Dabigatran Etexilate Mesylate Capsules 110mg Manufacturer in India. India has a well-established pharmaceutical manufacturing and distribution ecosystem, enabling manufacturers and suppliers to serve domestic distributors as well as international pharmaceutical markets. When sourcing an […]
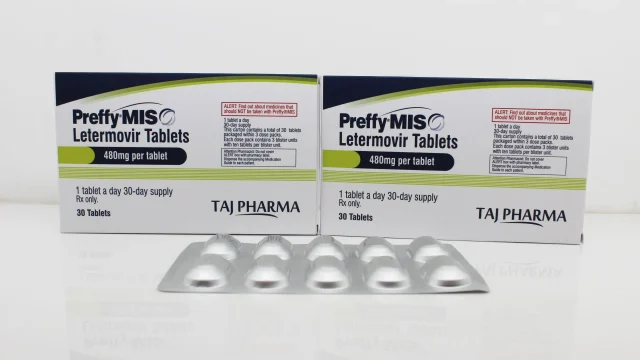
Letermovir Tablets 480mg Manufacturer and Supplier in India Taj Medico for Global Pharmaceutical Supply Taj Medico is an established pharmaceutical manufacturer, supplier, and exporter in India, offering pharmaceutical products for domestic and international markets. Among specialized antiviral products, Letermovir Tablets 480mg are associated with an important preventive role in patients at risk of cytomegalovirus (CMV) infection and disease following transplantation. Letermovir is an antiviral medicine specifically developed to prevent CMV reactivation and related disease in certain transplant recipients. It is particularly important in CMV-seropositive recipients of an allogeneic hematopoietic stem cell transplant (HSCT). Regulatory information confirms that letermovir acts by inhibiting the CMV terminase complex, an essential viral mechanism involved in the processing and packaging of viral DNA. For pharmaceutical buyers searching for a Letermovir 480mg manufacturer in India, Letermovir Tablets supplier, Letermovir 480mg exporter, or pharmaceutical manufacturing partner, Taj Medico provides a professional supply-oriented approach focused on quality, documentation, packaging, and international market requirements. Letermovir Tablets 480mg Manufacturer in India Letermovir Tablets 480mg We understand that specialized antiviral medicines require consistent manufacturing standards, reliable quality systems, and careful handling throughout the pharmaceutical supply chain. As a Letermovir Tablets 480mg manufacturer and supplier in India, we focus on maintaining appropriate […]
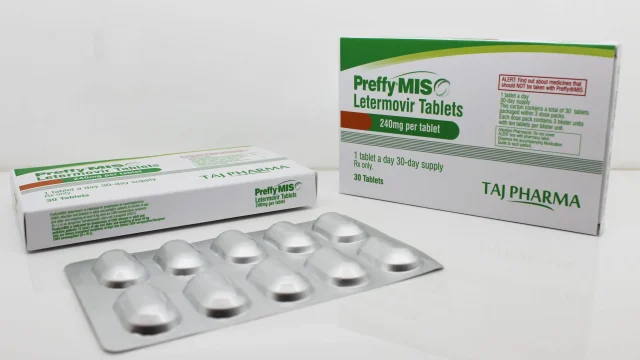
Letermovir 240mg Tablets Manufacturer & Supplier in India Taj Medico is a pharmaceutical manufacturer, exporter, and supplier based in Vapi, Gujarat, India, offering Letermovir 240mg Tablets for eligible domestic and international pharmaceutical markets. We focus on quality-oriented manufacturing, dependable supply, appropriate pharmaceutical documentation, and professional support for authorized buyers. Letermovir is an antiviral medicine used for cytomegalovirus (CMV) prophylaxis in specific patients, particularly certain adults who have undergone an allogeneic hematopoietic stem-cell transplant (HSCT) and are at increased risk of clinically significant CMV infection or disease. It is important to distinguish Letermovir from unrelated medicines sometimes incorrectly associated with its use. Letermovir is not a treatment for leprosy or dermatitis herpetiformis. Letermovir 240mg Tablets Manufacturer in India Letermovir 240mg Tablets We provide pharmaceutical supply solutions for buyers looking for a Letermovir 240mg Tablets Manufacturer in India. Letermovir belongs to a specialized class of antiviral medicines and acts by inhibiting the CMV terminase complex, a mechanism distinct from many older anti-CMV medicines. The 240mg tablet strength is particularly relevant because Letermovir dosing can vary according to the patient’s clinical circumstances and concomitant medications. The prescribing healthcare professional should determine the appropriate dosage and duration according to approved product information. For pharmaceutical […]
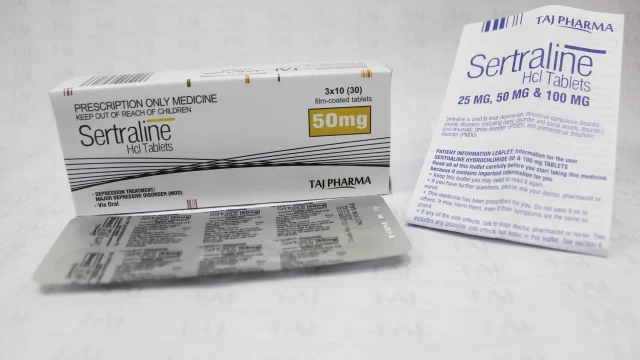
Sertraline Hcl Tablets 50mg PAN India Manufacturer Sertraline Hcl Tablets 50mg Manufacturer & Supplier in India We are a well-established Sertraline Hcl Tablets 50mg Manufacturer and Supplier in India, providing pharmaceutical products to distributors, wholesalers, institutional buyers, healthcare businesses, and international pharmaceutical partners. Based in Vapi, Gujarat, we focus on quality-oriented manufacturing, consistent supply, professional documentation, and export support. Sertraline Hcl Tablets 50mg contain sertraline hydrochloride, a selective serotonin reuptake inhibitor (SSRI) used as a prescription medicine for several approved mental health conditions. Depending on the applicable regulatory label, sertraline is used in the treatment of conditions such as major depressive disorder, obsessive-compulsive disorder, panic disorder, social anxiety disorder, and post-traumatic stress disorder. The 50mg tablet is an established strength used in pharmaceutical markets worldwide. We understand that buyers require more than a pharmaceutical product: they also need dependable manufacturing, appropriate quality documentation, secure packaging, responsive communication, and supply-chain coordination. Our approach is therefore designed around the complete requirements of pharmaceutical sourcing. Sertraline Hcl Tablets Manufacturer Suppliers in India Sertraline Hcl Tablets 50mg India has an important position in the global pharmaceutical supply chain because of its manufacturing capabilities, technical expertise, established regulatory infrastructure, and extensive export network. We serve […]
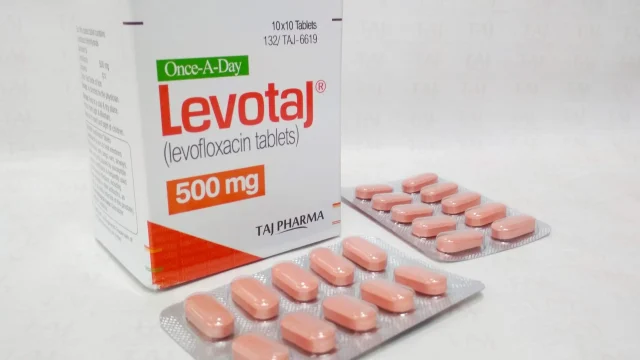
Levofloxacin 500mg Tablets Manufacturer Supplier in India – Taj Medico Levofloxacin 500mg Tablets Manufacturer in India Levofloxacin 500mg Tablets are prescription antibacterial medicines containing levofloxacin, a fluoroquinolone antibiotic used for selected bacterial infections caused by susceptible microorganisms. We at Taj Medico manufacture and supply pharmaceutical formulations from Vapi, Gujarat, India, supporting eligible distributors, importers, wholesalers, institutional buyers, and international pharmaceutical businesses. As a Levofloxacin 500mg Tablets manufacturer in India, we emphasize quality-focused production, consistent product specifications, organized supply, appropriate documentation, and professional customer support. Our objective is to provide a dependable sourcing option for pharmaceutical buyers requiring finished-dose levofloxacin products for approved markets. Levofloxacin should be prescribed only when clinically appropriate. It is not effective against viral infections, and treatment decisions should consider the causative organism, susceptibility, infection site, patient-specific factors, and applicable prescribing guidelines. What Are Levofloxacin 500mg Tablets? Levofloxacin Tablets Levofloxacin is a member of the fluoroquinolone class of antibacterial medicines. The 500mg tablet provides a specific oral strength that may be prescribed for certain susceptible bacterial infections. Levofloxacin works by interfering with bacterial enzymes involved in DNA replication and repair. Its principal antibacterial targets include DNA gyrase and topoisomerase IV, which are essential for bacterial DNA processes. […]
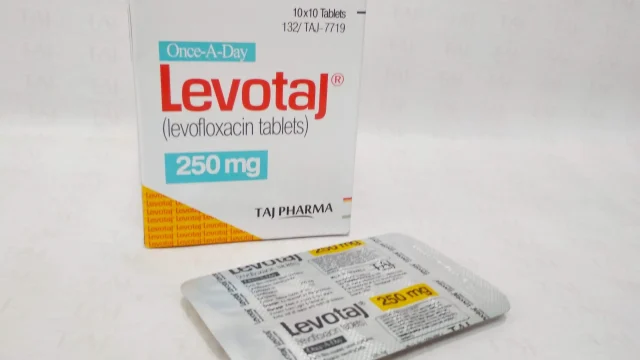
Levofloxacin Tablets 250mg Manufacturer Supplier in India – Taj Medico Levofloxacin Tablets 250mg are an important prescription antibiotic used for treating selected bacterial infections. At Taj Medico, we focus on the consistent manufacturing, quality assurance, and reliable supply of pharmaceutical products for domestic and international markets. As a Levofloxacin Tablets 250mg manufacturer and supplier in India, we support healthcare businesses, distributors, importers, wholesalers, and institutional buyers seeking dependable pharmaceutical sourcing from India. Levofloxacin belongs to the fluoroquinolone class of antibacterial medicines. It works by interfering with bacterial enzymes required for DNA replication and repair, thereby preventing susceptible bacteria from multiplying. Because resistance and patient-specific factors influence antibiotic selection, Levofloxacin should be supplied and used according to applicable medical, regulatory, and prescribing requirements. Levofloxacin Tablets 250mg Manufacturer in India Levofloxacin Tablets 250mg (Levotaj) We manufacture and supply Levofloxacin Tablets 250mg with an emphasis on product consistency, controlled production processes, quality testing, and appropriate pharmaceutical documentation. Our operations in Vapi, Gujarat, India, place us within one of India’s established pharmaceutical manufacturing regions and provide an efficient base for serving domestic and export markets. Our approach to pharmaceutical manufacturing centers on maintaining consistent product specifications from batch to batch. We understand that buyers […]
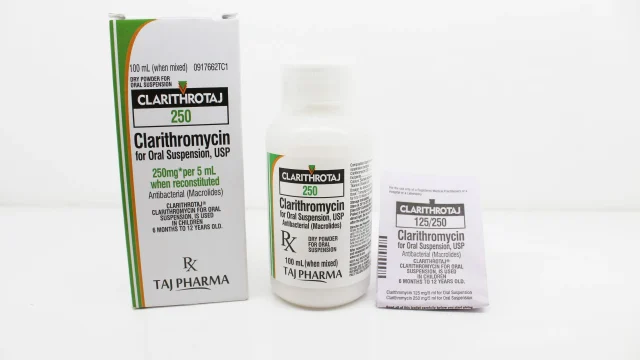
Clarithromycin for Suspension 250mg Manufacturer & Supplier in India Taj Medico is an established pharmaceutical manufacturer, exporter, and supplier based in Vapi, Gujarat, India, offering Clarithromycin for Suspension 250mg, marketed as Clarithrotaj, for eligible pharmaceutical distribution and procurement requirements. Clarithromycin is a macrolide antibiotic used against susceptible bacterial infections, including certain infections affecting the respiratory tract, skin and soft tissues, ear, throat, and other sites. As a pharmaceutical supplier, we focus on quality-oriented manufacturing, consistent product specifications, professional packaging, supply coordination, and responsive customer service. Our objective is to support pharmaceutical distributors, wholesalers, institutional buyers, eligible PCD pharma franchise partners, and international customers seeking dependable sourcing from India. ◆ Clarithromycin for Suspension 250mg Manufacturer in India. Clarithromycin for Suspension USP 250mg We supply Clarithromycin for Suspension 250mg to meet the requirements of pharmaceutical businesses looking for dependable products from an Indian manufacturer and supplier. Clarithromycin belongs to the macrolide class of antibiotics. It works primarily by interfering with bacterial protein synthesis, preventing susceptible bacteria from producing proteins needed for growth and reproduction. The 250mg oral suspension provides Clarithromycin in a liquid dosage form. This presentation can be useful when an oral liquid formulation is preferred, including situations where swallowing solid […]
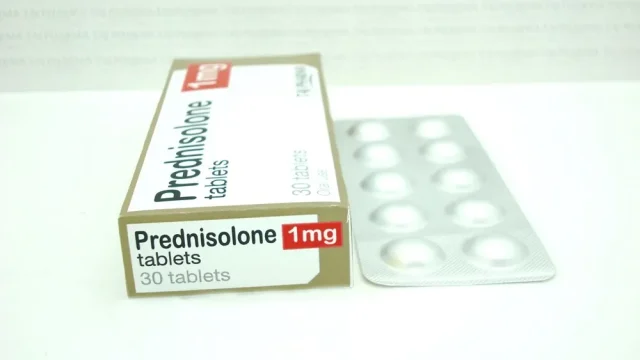
Prednisolone 1mg Tablets Manufacturer and Supplier in India Prednisolone 1mg Tablets Manufacturer in India We at Taj Medico focus on supplying high-quality pharmaceutical products to domestic and international markets, including Prednisolone 1mg Tablets. As a manufacturer, supplier, and exporter based in Gujarat, India, we work to support pharmaceutical distributors, importers, wholesalers, hospitals, institutional buyers, and other authorized healthcare-sector partners seeking reliable finished dosage formulations. Prednisolone is a corticosteroid medicine used under the supervision and prescription of qualified healthcare professionals for a range of inflammatory, allergic, autoimmune, and other medical conditions. The medicine works by modifying immune and inflammatory responses in the body. Because corticosteroids can have significant effects and may cause adverse reactions, Prednisolone should always be used according to the advice of a qualified medical professional. With growing demand for dependable pharmaceutical sourcing across India, Africa, the Middle East, Latin America, and other international markets, we understand that buyers require more than a product alone. They need consistency, professional communication, appropriate documentation, dependable supply capabilities, and packaging options suited to their market requirements. Our objective is to build long-term business relationships by supporting these requirements throughout the sourcing and supply process. About Prednisolone 1mg Tablets Prednisilone 1mg Tablets Prednisolone […]
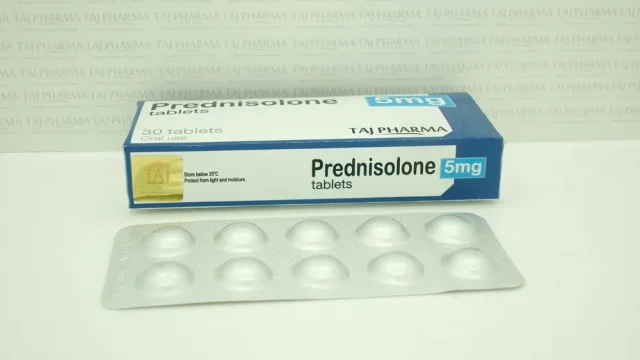
Prednisolone 5mg Tablets Manufacturer and Supplier in India Prednisolone 5mg Tablets Manufacturer in India At Taj Medico, we are committed to supplying high-quality pharmaceutical products for domestic and international markets. As a Prednisolone 5mg Tablets manufacturer and supplier in India, we focus on product quality, consistent manufacturing standards, reliable supply, and professional support for pharmaceutical distributors, importers, wholesalers, healthcare procurement companies, and other eligible business partners. Prednisolone 5mg Tablets are corticosteroid medicines used under appropriate medical supervision for conditions where reducing inflammation or modifying immune-system activity is clinically required. Prednisolone belongs to the corticosteroid class of medicines and may be prescribed for a broad range of medical conditions, depending on individual patient needs and the judgement of a qualified healthcare professional. We understand that pharmaceutical buyers require more than a competitive quotation. They also look for dependable manufacturing processes, suitable packaging, clear product information, export coordination, and consistent availability. Our approach is designed to support these important requirements while serving customers across India, Africa, the Middle East, Latin America, and other international markets. Reliable Prednisolone 5mg Tablets Supplier and Exporter Prednisilone 5mg Tablets We serve as a reliable source for businesses seeking Prednisolone Tablets from India. Our supply capabilities are […]
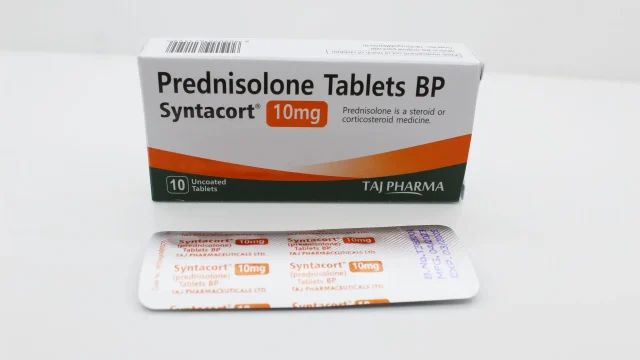
Prednisolone BP 10mg Tablets Manufacturer & Supplier in India – Taj Medico for Global Pharmaceutical Buyers We are Taj Medico, an established pharmaceutical manufacturer, exporter, and supplier based in Vapi, Gujarat, India, offering Prednisolone BP 10mg Tablets for pharmaceutical distributors, importers, wholesalers, healthcare businesses, and international buyers. We focus on consistent manufacturing standards, reliable supply, quality-oriented processes, and professional export support for customers seeking dependable finished pharmaceutical products. Our Prednisolone BP 10mg Tablets are formulated with prednisolone, a corticosteroid medicine that is used under medical supervision for a range of inflammatory, allergic, immune-related, and other clinical conditions. As a manufacturer and exporter, we work to provide commercially reliable pharmaceutical solutions while supporting the documentation and supply requirements of domestic and international markets. Prednisolone BP 10mg Tablets Manufacturer in India Prednisolone Tablets 10mg We manufacture and supply Prednisolone BP 10mg Tablets for buyers looking for a dependable Indian pharmaceutical partner. Our location in Vapi, Gujarat, places us within one of India’s important pharmaceutical manufacturing regions, providing access to established pharmaceutical infrastructure, skilled professionals, packaging capabilities, and supply-chain networks. We understand that pharmaceutical buyers require more than a product name and competitive pricing. They need consistency, dependable communication, appropriate documentation, professional packaging, […]
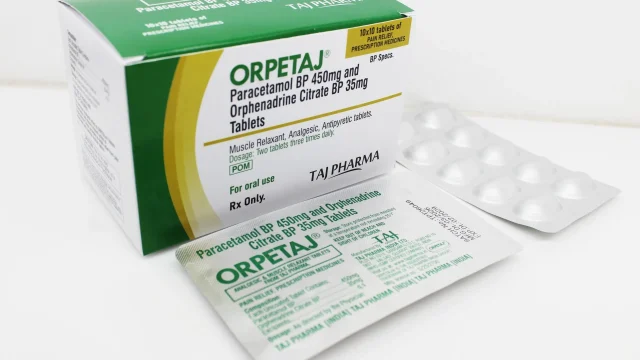
Paracetamol 450mg and Orphenadrine Citrate 35mg Tablets Manufacturer & Supplier in India Leading Manufacturer, Supplier and PCD Pharma Franchise Provider for Tablets We at Taj Medico are committed to serving the pharmaceutical market with reliable manufacturing, supply, distribution, and business partnership solutions for quality-focused pharmaceutical products. Paracetamol 450mg and Orphenadrine Citrate 35mg Tablets, marketed under the brand name ORPETAJ, are an important product offering for pharmaceutical distributors, wholesalers, healthcare supply businesses, and PCD Pharma Franchise partners seeking a dependable source from India. As a Paracetamol and Orphenadrine Tablets manufacturer and supplier, we focus on maintaining consistency in product handling, packaging, supply coordination, and commercial support. Our objective is to build long-term relationships with customers across Mumbai, Maharashtra, Gujarat, Vapi, and PAN India, while also supporting international business requirements where applicable. With growing demand for professionally managed pharmaceutical supply chains, buyers increasingly look for manufacturers and suppliers who can provide responsive communication, dependable order coordination, and scalable business opportunities. Through ORPETAJ Tablets, we aim to provide a structured solution for businesses searching for Paracetamol 450mg and Orphenadrine Citrate 35mg Tablets manufacturers in India. ORPETAJ – Paracetamol 450mg and Orphenadrine Citrate 35mg Tablets Paracetamol 450mg and Orphenadrine Citrate 35mg Tablets ORPETAJ contains: […]
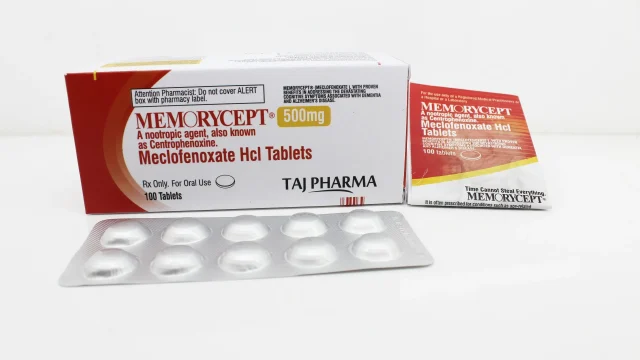
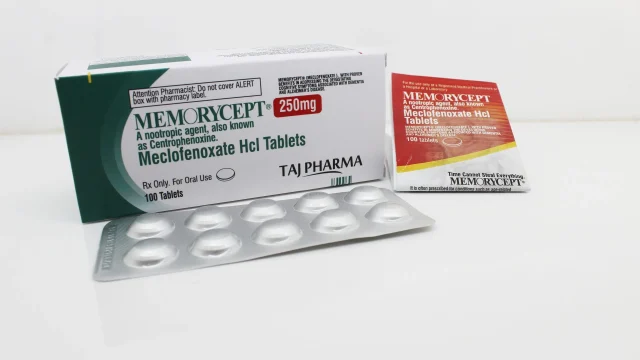
Meclofenoxate Tablets 500mg Manufacturer Supplier in India Meclofenoxate Tablets 500mg Manufacturer in India Taj Medico is an established Meclofenoxate Tablets 500mg Manufacturer and Supplier in India, serving qualified pharmaceutical wholesalers, distributors, importers, and authorized healthcare businesses. Based in Vapi, Gujarat, we support PAN India requirements and eligible international pharmaceutical markets through organized supply coordination, professional communication, and market-specific procurement support. Meclofenoxate, also known as centrophenoxine, is a compound classified in pharmaceutical references under nootropic and neuroprotective agents. It is associated with the nervous system rather than immunosuppression or pain relief. PubChem lists meclofenoxate under the WHO ATC category N06BX01 and identifies centrophenoxine as a synonym. This distinction is important because some online descriptions incorrectly identify meclofenoxate as an immunosuppressant or NSAID. Meclofenoxate should not be confused with medicines used to prevent organ-transplant rejection or with analgesic anti-inflammatory medicines. Meclofenoxate Tablets 500mg Supplier for Pharmaceutical Buyers Meclofenoxate Tablets 500mg As a professional Meclofenoxate Tablets 500mg Supplier, we understand the importance of dependable pharmaceutical procurement. Buyers require more than a product name and strength; they also need appropriate specifications, packaging, batch information, documentation, logistics planning, and regulatory clarity. We work with qualified pharmaceutical businesses seeking: Bulk pharmaceutical supply Wholesale procurement PAN India […]
Meclofenoxate Tablets 250mg Manufacturer Supplier in India Meclofenoxate Tablets 250mg Manufacturer in India Taj Medico is an established Meclofenoxate Tablets 250mg Manufacturer and Supplier in India, providing pharmaceutical sourcing solutions for qualified wholesalers, distributors, importers, and authorized healthcare businesses. Based in Vapi, Gujarat, we support PAN India requirements and eligible international markets through organized supply coordination, professional communication, and customer-focused service. Meclofenoxate Tablets 250mg contain meclofenoxate, also known as centrophenoxine, an active pharmaceutical compound classified by the National Library of Medicine under nootropic and neuroprotective agents. It is not an immunosuppressant; claims that meclofenoxate prevents transplanted-organ rejection or suppresses the immune system are inaccurate. Meclofenoxate has been studied in relation to cognitive function and neurocognitive disorders, although approved indications and availability can differ between countries. Buyers should therefore verify the product’s regulatory status and approved uses in their intended market before procurement. Meclofenoxate Tablets 250mg Supplier for Global Pharmaceutical Buyers Meclofenoxate Tablets 250mg As a professional Meclofenoxate Tablets 250mg Supplier, we understand that pharmaceutical procurement requires more than product availability. Importers and distributors need dependable communication, appropriate product specifications, suitable packaging, accurate batch information, documentation, and coordinated logistics. We work with qualified pharmaceutical businesses seeking: Bulk pharmaceutical procurement Wholesale supply […]
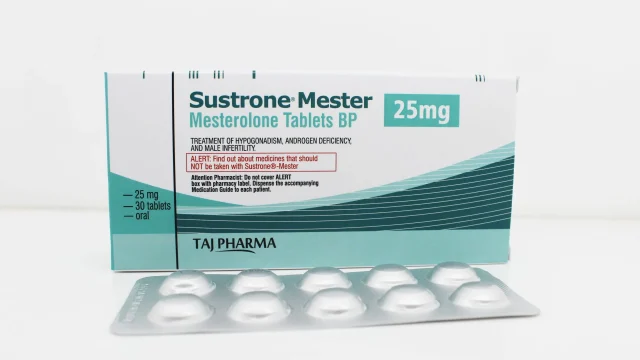
Mesterolone BP 25mg Tablets Manufacturer and Supplier in India Leading Mesterolone BP 25mg Tablets Manufacturer in India We at Taj Medico are committed to supplying high-quality pharmaceutical products for domestic and international markets. As an established Mesterolone BP 25mg Tablets manufacturer, supplier, and exporter in India, we focus on consistent product quality, professional service, reliable documentation, and dependable supply capabilities. Mesterolone BP 25mg Tablets are pharmaceutical preparations containing mesterolone, an androgenic medicine used under appropriate medical supervision in the management of certain conditions associated with androgen deficiency, including male hypogonadism. Because hormonal medicines require careful prescribing and monitoring, responsible manufacturing, accurate labeling, controlled distribution, and regulatory compliance are essential throughout the supply chain. From our operations in Vapi, Gujarat, India, we serve pharmaceutical buyers, distributors, wholesalers, importers, institutional customers, and international business partners seeking a dependable source for Mesterolone 25mg Tablets. Our approach is centered on maintaining consistency from procurement and manufacturing through packaging, quality evaluation, documentation, dispatch, and export coordination. Mesterolone BP 25mg Tablets – Product Overview Mesterolone Tablets BP 25mg Mesterolone is an orally administered androgenic compound. It may be prescribed by qualified healthcare professionals for specific medical indications involving insufficient androgen activity. Treatment decisions, dosage, duration, and […]
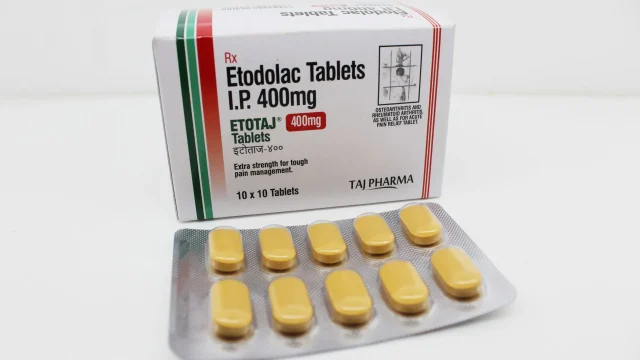
Etodolac Tablets 400mg Manufacturer, Supplier & Exporter in India Etodolac Tablets 400mg Manufacturer in India We, at Taj Medico, are an established manufacturer, supplier, and exporter of Etodolac Tablets 400mg (Etotaj) in India, serving pharmaceutical businesses, distributors, wholesalers, healthcare organizations, and international buyers. From our operations in Vapi, Gujarat, we focus on supplying pharmaceutical products that meet defined quality standards and are supported by consistent manufacturing and supply practices. Etodolac Tablets 400mg contain etodolac, a non-steroidal anti-inflammatory drug (NSAID) used for the management of pain and inflammatory symptoms associated with conditions such as osteoarthritis and rheumatoid arthritis. Depending on the patient’s condition and the prescribing physician’s assessment, etodolac may help reduce pain, swelling, tenderness, and stiffness. As a dedicated Etodolac Tablets 400mg Manufacturer in India, we understand that pharmaceutical buyers require more than product availability. They also need dependable quality, suitable packaging, regulatory documentation, competitive commercial terms, and reliable order fulfillment. We therefore focus on building long-term supply relationships with domestic and international pharmaceutical partners. Etodolac Tablets 400mg – Product Overview Etodolac Tablets 400mg (Etotaj) Etodolac belongs to the NSAID category and is primarily used for its analgesic and anti-inflammatory properties. It works by reducing the production of prostaglandins, chemical […]
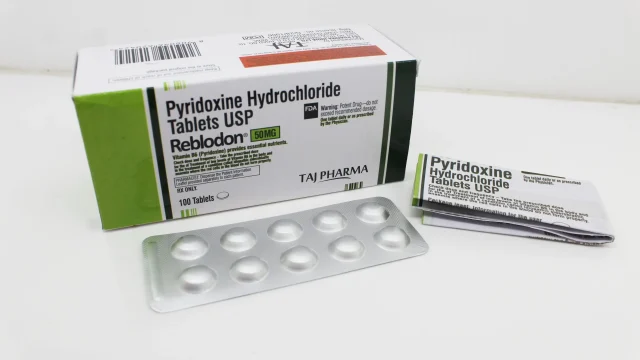
Pyridoxine Hydrochloride 50mg Tablets Manufacturer and Supplier in India Taj Medico is a recognized Pyridoxine Hydrochloride 50mg Tablets manufacturer, supplier, and exporter in India, supporting pharmaceutical distributors, wholesalers, importers, and authorized healthcare procurement organizations. From Vapi, Gujarat, we work with customers across PAN India and international markets, including Africa, the Middle East, Latin America, and other regions, subject to applicable regulatory and import requirements. Our Pyridoxine Hydrochloride 50mg Tablets, also marketed under the brand name Reblodon, are formulated to provide pyridoxine, commonly known as Vitamin B6. Vitamin B6 is an essential nutrient involved in numerous metabolic and physiological functions. Pharmaceutical-grade Pyridoxine Hydrochloride formulations may be prescribed for the prevention or treatment of vitamin B6 deficiency and other medically appropriate indications. As a dependable Pyridoxine Hydrochloride 50mg Tablets manufacturer in India, we focus on professional pharmaceutical supply, quality-oriented manufacturing practices, appropriate packaging, dependable communication, and support for qualified domestic and international buyers. Pyridoxine Hydrochloride 50mg Tablets – Product Overview Pyridoxine Hydrochloride 50mg Tablets Pyridoxine Hydrochloride is a form of Vitamin B6 used in pharmaceutical and nutritional formulations. Vitamin B6 plays an important role in amino acid metabolism, neurotransmitter synthesis, hemoglobin formation, and other biological processes. Pyridoxine Hydrochloride 50mg Tablets may be […]
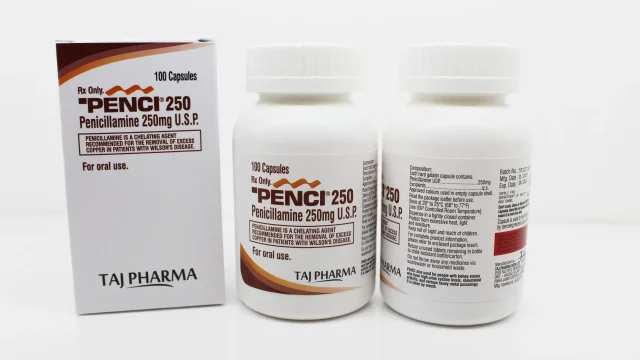
Pencillamine 250mg Capsules Manufacturer and Supplier in India – Trusted Global Exporter Pencillamine 250mg Capsules are an important prescription pharmaceutical formulation used in the management of specific medical conditions, including Wilson’s disease, cystinuria, and selected cases of severe active rheumatoid arthritis. As international pharmaceutical markets continue to demand dependable sourcing, consistent quality standards, and professional export support, choosing an experienced Pencillamine 250mg Capsules manufacturer becomes increasingly important. At Taj Medico, we focus on supplying high-quality pharmaceutical products to domestic and international buyers. As a recognized Pencillamine 250mg Capsules manufacturer, supplier, and exporter from Vapi, Gujarat, India, we support pharmaceutical distributors, wholesalers, importers, healthcare procurement companies, and other authorized business partners seeking reliable pharmaceutical supply solutions. Our objective is to provide professionally manufactured pharmaceutical products supported by consistent quality practices, appropriate packaging, dependable supply capabilities, and export-oriented service for customers across India, Africa, the Middle East, Latin America, and other international markets. Pencillamine 250mg Capsules – Product Overview Penicillamine 250mg Capsules Pencillamine, also known as D-Penicillamine, is a prescription medicine with chelating and immunomodulatory properties. It has established therapeutic applications in condiions where reducing or binding certain substances in the body is medically necessary. Pencillamine can bind with substances such as […]
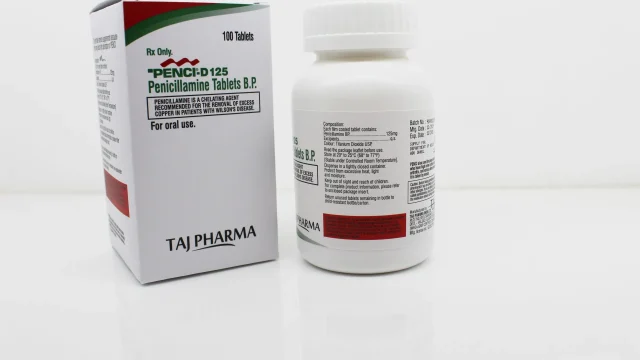
Penicillamine BP 125mg Tablets Manufacturer in India – Taj Medico for Global Pharmaceutical Supply We are Taj Medico, an established Penicillamine BP 125mg Tablets Manufacturer, Supplier, and Exporter in India, committed to supplying pharmaceutical products with consistent quality, reliable manufacturing standards, and professional documentation. Our Penicillamine 125mg Tablets (Penici-D) are positioned for pharmaceutical distributors, institutional buyers, importers, wholesalers, and healthcare markets seeking a dependable Indian manufacturing and supply partner. Penicillamine is a prescription medicine with important applications in Wilson’s disease, cystinuria, and selected cases of severe active rheumatoid arthritis. Because the medicine requires careful patient selection, monitoring, and dose adjustment, responsible pharmaceutical manufacturing and supply are particularly important. We focus on maintaining appropriate quality controls throughout manufacturing, packaging, storage, and distribution. Penicillamine BP 125mg Tablets Manufacturer in India Penicillamine-D 125mg Tablets As a Penicillamine BP 125mg Tablets Manufacturer in India, we understand that pharmaceutical buyers require more than product availability. They need dependable quality, consistent specifications, appropriate packaging, documentation, and responsive supply support. Our manufacturing operations are based in Vapi, Gujarat, India, a major pharmaceutical manufacturing region. We work toward maintaining controlled production processes designed to support batch-to-batch consistency. Our approach incorporates quality-oriented manufacturing practices, suitable raw-material controls, in-process checks, […]
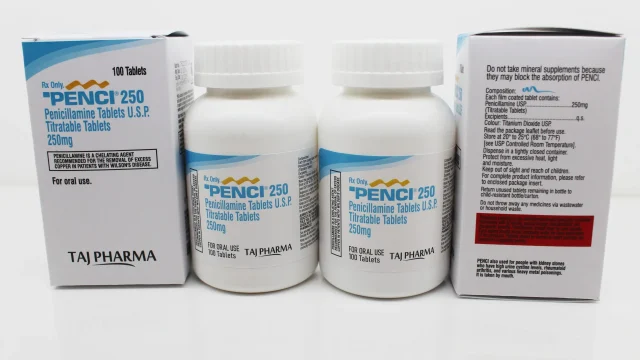
Penicillamine USP 250mg Manufacturer, Supplier and Exporter in India Taj Medico is an established Penicillamine USP 250mg Manufacturer, Supplier and Exporter in India, offering pharmaceutical supply solutions from Vapi, Gujarat. We focus on quality-oriented manufacturing, consistent product specifications, professional documentation, and dependable supply for authorized pharmaceutical businesses across India and international markets. Penicillamine 250mg Tablets (Penci) 🏭 Penicillamine USP 250mg Manufacturer in India We manufacture and supply Penicillamine USP 250mg Tablets for pharmaceutical distributors, wholesalers, institutional buyers, exporters, and other authorized healthcare businesses. Our manufacturing approach emphasizes controlled production, quality assurance, finished-product testing, appropriate packaging, and reliable batch-to-batch consistency. Penicillamine is a specialized prescription medicine with chelating and pharmacological properties. Because its use can require regular clinical and laboratory monitoring, pharmaceutical supply must be supported by appropriate quality systems and responsible distribution. 📍 Manufacturing Location: Vapi, Gujarat, India💊 Product: Penicillamine Tablets⚖️ Strength: 250mg📋 Standard: USP🌍 Supply: PAN India and export markets🏢 Business Type: Manufacturer, Supplier and Exporter 💊 Penicillamine 250mg Tablets Uses Penicillamine 250mg has established medical applications in several specific conditions. 🧬 1. Wilson’s Disease Penicillamine is used as a copper-chelating medicine in Wilson’s disease. This inherited disorder causes excessive copper accumulation in tissues. Penicillamine binds copper and facilitates […]
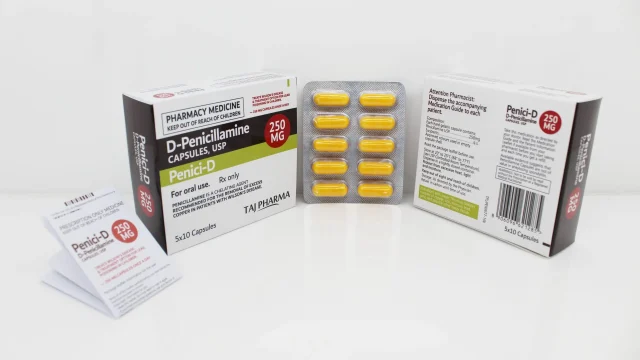
D-Penicillamine USP 250mg Capsules Manufacturer & Supplier in India Taj Medico is an established pharmaceutical manufacturer and supplier associated with D-Penicillamine USP 250mg Capsules (Penici-D) for authorized pharmaceutical procurement in India and eligible international markets. We focus on providing consistent pharmaceutical products, dependable supply coordination, and professional support to distributors, wholesalers, importers, healthcare procurement organizations, and other authorized buyers. D-Penicillamine 250mg Capsules contain D-penicillamine, a prescription medicine with chelating and immunomodulatory properties. Depending on the clinical indication, D-penicillamine may be prescribed for Wilson’s disease, severe active rheumatoid arthritis, and cystinuria. Because this medicine can cause important adverse effects and generally requires laboratory monitoring, treatment should always be directed by a qualified healthcare professional. Our objective is to support legitimate pharmaceutical supply requirements with appropriate product information, quality-focused manufacturing practices, packaging coordination, and documentation according to applicable market requirements. D-Penicillamine USP 250mg Capsules – Product Overview D- Penicillamine USP 250mg Capsules D-Penicillamine USP 250mg Capsules, marketed as Penici-D, are prescription capsules containing 250 mg of D-penicillamine. D-penicillamine is a thiol-containing compound with two important pharmacological characteristics. First, it has chelating activity, allowing it to bind certain substances and facilitate their elimination. Second, it has immunomodulatory activity, which contributes to its historical […]
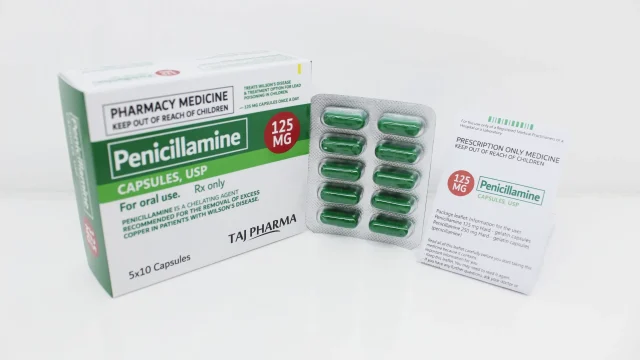
Penicillamine Capsules 125mg Manufacturer, Supplier & Exporter in India Taj Medico is a well-established Penicillamine Capsules 125mg manufacturer, supplier, and exporter in India, offering pharmaceutical products for domestic and international markets. We focus on consistent manufacturing standards, reliable quality, appropriate packaging, and dependable supply support for healthcare businesses and authorized pharmaceutical distributors. Penicillamine Capsules 125mg contain penicillamine, a prescription medicine with chelating and disease-modifying properties. Depending on the clinical indication, penicillamine may be prescribed for Wilson’s disease, severe rheumatoid arthritis, and cystinuria. Because penicillamine can cause significant adverse reactions and requires laboratory monitoring, treatment should be initiated and supervised by a qualified healthcare professional. Our focus as a pharmaceutical supplier is to support authorized buyers with professionally manufactured products, suitable documentation, quality-focused processes, and dependable commercial supply. Penicillamine Capsules 125mg Manufacturer in India Penicillamine Capsules As a Penicillamine Capsules 125mg manufacturer in India, we understand that pharmaceutical buyers require more than product availability. They need consistency, traceability, appropriate packaging, documentation, and dependable communication throughout the procurement process. At Taj Medico, we work toward maintaining controlled manufacturing and quality-assurance practices across the product lifecycle. Our approach is designed to support pharmaceutical distributors, institutional buyers, importers, and other authorized organizations seeking reliable […]
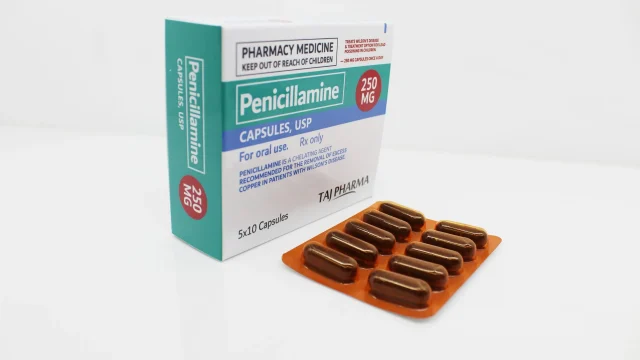
Penicillamine USP 250mg Manufacturer and Supplier in India Taj Medico is an established pharmaceutical manufacturer, supplier, and exporter of Penicillamine USP 250mg Capsules from Vapi, Gujarat, India. Penicillamine is a prescription medicine with chelating and disease-modifying properties. It is used under specialist medical supervision for conditions including Wilson’s disease, cystinuria, and severe active rheumatoid arthritis that has not responded adequately to conventional treatment. With an emphasis on pharmaceutical quality, controlled manufacturing, consistent product specifications, and supply reliability, we support domestic and international requirements for Penicillamine USP 250mg. Our approach focuses on maintaining appropriate quality systems throughout manufacturing, packaging, documentation, and supply. For buyers searching for Penicillamine 250mg Capsules Manufacturers in India, Penicillamine USP 250mg Suppliers, Penicillamine Manufacturers and Exporters, or Penicillamine 250mg pharmaceutical suppliers, Taj Medico provides a focused manufacturing and export solution. Penicillamine USP 250mg Capsules – Product Overview Penicillamine USP 250mg Capsules Penicillamine is a thiol-containing compound with an important role in chelation therapy. In Wilson’s disease, it binds copper and facilitates its elimination, helping reduce excessive copper accumulation. In cystinuria, it forms a more soluble complex with cystine, reducing cystine-related stone formation. For selected patients with severe active rheumatoid arthritis, penicillamine has disease-modifying activity and may be […]
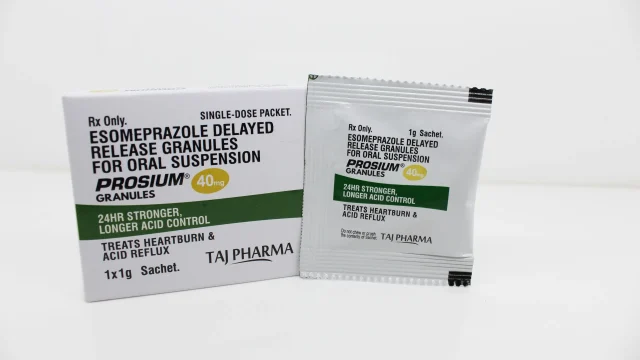
Esomeprazole Sachet Suspension 40mg Manufacturer in India – Premium Exporter & Global Supplier | Taj Medico Esomeprazole Sachet Suspension 40mg Manufacturer in India Taj Medico is a trusted Esomeprazole Sachet Suspension 40mg Manufacturer in India, committed to delivering high-quality pharmaceutical formulations that meet international quality standards. We manufacture and export premium-grade Esomeprazole Sachet Suspension 40mg for domestic and global pharmaceutical markets with a strong focus on quality, consistency, and regulatory compliance. Our manufacturing facility in Vapi, Gujarat, India follows stringent Good Manufacturing Practices (GMP), ensuring every batch is produced under controlled conditions for maximum safety, efficacy, and stability. We supply Esomeprazole Sachet Suspension 40mg across India, Africa, the Middle East, Latin America (LATAM), Asia, and other international markets, making us a preferred pharmaceutical manufacturing partner for distributors, importers, hospitals, and healthcare organizations. Esomeprazole Sachet Suspension 40mg What is Esomeprazole Sachet Suspension 40mg? Esomeprazole Sachet Suspension 40mg is a proton pump inhibitor (PPI) that effectively reduces the production of stomach acid. It is widely prescribed for treating various acid-related gastrointestinal disorders, providing long-lasting relief from excessive stomach acid and promoting healing of damaged tissues. The sachet formulation is particularly suitable for patients who have difficulty swallowing tablets, offering convenient administration while […]
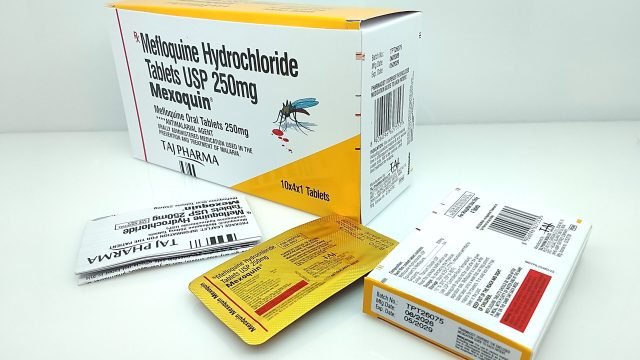
Mefloquine Hydrochloride Tablets 250mg (Mexoquin) Manufacturer, Supplier & Exporter in India Trusted Mefloquine Hydrochloride Tablets 250mg Manufacturer in India Taj Medico is a trusted manufacturer, supplier, and exporter of Mefloquine Hydrochloride Tablets 250mg (Mexoquin) in India. We are committed to delivering high-quality pharmaceutical products manufactured under stringent quality management systems using carefully selected raw materials and advanced manufacturing processes. Our production facilities are designed to maintain consistent quality, product integrity, and batch-to-batch reliability for domestic and international markets. We supply Mefloquine Hydrochloride Tablets 250mg to pharmaceutical distributors, hospitals, healthcare institutions, government organizations, wholesalers, retailers, and international importers across India, Africa, the Middle East, Latin America (LATAM), and other global markets. About Mefloquine Hydrochloride Tablets 250mg Mefloquine Hydrochloride Tablets 250mg Mefloquine Hydrochloride Tablets 250mg is a prescription antimalarial medicine indicated for the prevention and treatment of certain malaria infections when prescribed by a qualified healthcare professional. The medicine acts against malaria parasites and should be used only according to the advice of a healthcare provider and applicable treatment guidelines. For malaria prevention in travelers, dosing recommendations depend on individual circumstances and current medical guidance. Patients should consult their physician before starting therapy. Quality Manufacturing Standards At Taj Medico, every batch of […]
Fenbendazole Tablets 222mg Manufacturer in India – Premium Supplier & Exporter Fenbendazole Tablets 222mg have become one of the most trusted veterinary anthelmintic medicines for the treatment and control of a broad range of gastrointestinal parasites in animals. As a leading Fenbendazole Tablets 222mg Manufacturer in India, we at Taj Medico are committed to delivering premium-quality veterinary pharmaceutical products that meet international quality standards. Our advanced manufacturing practices, strict quality control procedures, and reliable export capabilities have made us a preferred partner for distributors, veterinary hospitals, wholesalers, importers, and government procurement agencies across Africa, the Middle East, LATAM, and numerous international markets. Manufactured at our modern facility in Vapi, Gujarat, India, our Fenbendazole Tablets 222mg (Fencure) are produced using carefully selected raw materials and validated manufacturing processes to ensure consistent quality, purity, stability, and effectiveness. We continue to strengthen our position as a reliable Fenbendazole Tablets Manufacturer, Supplier, and Exporter by delivering products that comply with global pharmaceutical manufacturing standards. Fenbendazole Tablets 222mg Manufacturer in India Fenbendazole Tablets 222mg We manufacture Fenbendazole Tablets 222mg specifically for veterinary use. Fenbendazole belongs to the benzimidazole class of anthelmintic medicines and is widely used to control a broad spectrum of gastrointestinal parasites in […]
Doxepin Hydrochloride Capsules 25mg Manufacturer in India – Trusted Global Supplier & Exporter by Taj Medico Doxepin Hydrochloride Capsules 25mg is a widely prescribed tricyclic antidepressant used for the treatment of major depressive disorder, anxiety disorders, chronic insomnia, and pruritus (skin itching). As demand for high-quality psychiatric medicines continues to grow across global healthcare markets, selecting a reliable pharmaceutical manufacturing partner becomes essential. Taj Medico, based in Vapi, Gujarat, India, is a leading Manufacturer, Exporter, and Supplier of Doxepin Hydrochloride Capsules 25mg (Doxphoria). We manufacture premium-quality formulations in compliance with international quality standards and serve pharmaceutical importers, government tenders, hospitals, distributors, wholesalers, and PCD franchise companies throughout India, Africa, the Middle East, LATAM, Southeast Asia, and other international markets. Our commitment to quality manufacturing, regulatory compliance, competitive pricing, and timely deliveries has established Taj Medico as a trusted pharmaceutical export partner for Doxepin Hydrochloride Capsules worldwide. Doxepin Hydrochloride Capsules 25mg – Product Overview Doxepin Hydrochloride Capsules I.P. 25mg Doxepin Hydrochloride belongs to the class of tricyclic antidepressants (TCAs). It acts by increasing the availability of neurotransmitters such as serotonin and norepinephrine in the brain, helping improve mood, reduce anxiety, and promote healthy sleep patterns. At the 25mg dosage, Doxepin is […]
Propofol 1% 10mg (Diprikoda10) Manufacturer Supplier in India – Trusted Quality from Taj Medico Leading Propofol 1% 10mg Manufacturer in India Taj Medico is a trusted Propofol 1% 10mg (Diprikoda10) Manufacturer, Supplier, and Exporter in India, delivering premium-quality sterile injectable formulations to hospitals, healthcare institutions, pharmaceutical distributors, and international buyers. Manufactured using advanced pharmaceutical technology and stringent quality control systems, our Propofol injection is supplied across PAN India and exported to customers in Africa, the Middle East, LATAM, and other global markets. Our commitment to pharmaceutical excellence, reliable manufacturing, and customer satisfaction has established Taj Medico as a dependable partner for institutional procurement, government tenders, pharmaceutical wholesalers, and healthcare organizations seeking consistent quality and timely deliveries. What is Propofol 1% 10mg? Propofol 1% 10mg Propofol 1% 10mg is an intravenous anesthetic medicine administered by qualified healthcare professionals for: Induction of general anesthesia Maintenance of general anesthesia Procedural sedation Sedation in Intensive Care Units (ICUs) Propofol has a rapid onset of action and is widely used in hospitals and surgical centers where controlled anesthesia and sedation are required. Because it is a potent anesthetic agent, it must always be administered under the supervision of trained medical professionals in an appropriate clinical […]
Chlorpheniramine Maleate Tablets 4mg Manufacturer Supplier in India – Trusted PAN India Exporter, PCD Pharma Franchise & Global Pharmaceutical Partner Leading Chlorpheniramine Maleate Tablets 4mg Manufacturer in India Taj Medico is a leading Chlorpheniramine Maleate Tablets 4mg manufacturer, supplier, exporter, and PCD Pharma Franchise company in India, delivering premium-quality pharmaceutical formulations that comply with stringent international manufacturing standards. Manufactured at our advanced facility in Vapi, Gujarat, our (Pollentaj) Chlorpheniramine Maleate Tablets 4mg are produced using high-quality Active Pharmaceutical Ingredients (APIs) under strict WHO-GMP quality systems to ensure safety, efficacy, and batch-to-batch consistency. We serve pharmaceutical distributors, hospitals, wholesalers, retailers, government institutions, healthcare organizations, and international importers with dependable manufacturing capabilities, competitive pricing, and reliable delivery schedules. About Chlorpheniramine Maleate Tablets 4mg Chlorpheniramine maleate Tablets 4mg Chlorpheniramine Maleate Tablets 4mg are an antihistamine medicine commonly prescribed for the symptomatic relief of allergic conditions. The medicine helps reduce the effects of histamine, a naturally occurring substance responsible for allergy symptoms. These tablets are widely recommended for managing: Allergic rhinitis Hay fever Skin allergies Urticaria (hives) Itching Sneezing Runny nose Watery eyes Insect bite reactions Seasonal allergies The tablets should always be taken according to the advice of a qualified healthcare professional. Premium […]
Paracetamol Injection 0.5% w/v Manufacturer in India – Trusted Global Supplier & Exporter | Taj Medico Paracetamol Injection 0.5% w/v is one of the most widely used intravenous analgesic and antipyretic medicines in hospitals and healthcare institutions worldwide. It is extensively administered for the short-term treatment of moderate to severe pain and for the reduction of fever when oral administration is not possible. As a trusted Paracetamol Injection 0.5% w/v Manufacturer in India, we at Taj Medico are committed to delivering pharmaceutical products that meet the highest international quality standards while ensuring consistent efficacy, safety, and regulatory compliance. With advanced manufacturing facilities in Vapi, Gujarat, India, we manufacture and export premium-quality Parataj I.V. (Paracetamol Injection 0.5% w/v) to hospitals, government institutions, pharmaceutical distributors, importers, and healthcare organizations across Africa, the Middle East, Latin America, Asia, and other international markets. Paracetamol Injection 0.5% w/v – Product Overview Paracetamol Injection 0.5 w/v Paracetamol Injection 0.5% w/v is a sterile intravenous solution formulated for rapid pain relief and fever management. It is particularly useful in clinical settings where oral medication cannot be administered due to surgery, unconsciousness, vomiting, or gastrointestinal conditions. Paracetamol works by inhibiting the production of prostaglandins in the central nervous […]
Paracetamol Injection 1% w/v Manufacturer in India – Trusted Supplier & Exporter | Taj Medico Paracetamol Injection 1% w/v is one of the most widely used intravenous analgesic and antipyretic formulations in modern healthcare. It is extensively administered in hospitals, emergency departments, intensive care units, and surgical centers for the effective management of pain and fever when oral administration is not suitable. As demand for high-quality injectable medicines continues to grow globally, healthcare providers and pharmaceutical distributors require a manufacturing partner that consistently delivers quality, safety, and reliability. At Taj Medico, we are a trusted Manufacturer, Exporter, and Supplier of Paracetamol Injection 1% w/v (Parataj I.V.) from Vapi, Gujarat, India. We manufacture pharmaceutical products in WHO-GMP compliant facilities using advanced technology, pharmaceutical-grade raw materials, and stringent quality control procedures. Our products are supplied throughout India and exported to Africa, the Middle East, LATAM, and many other international markets. Leading Paracetamol Injection 1% w/v Manufacturer in India Paracetamol Injection 1% w/v We manufacture Parataj I.V. under strict pharmaceutical quality systems to ensure every batch delivers consistent efficacy, sterility, and safety. Our manufacturing infrastructure is designed to comply with international pharmaceutical standards, making our injectable formulations suitable for both domestic and export […]
Miltefosine Capsules 50mg Manufacturer Supplier in India – Why Taj Medico is the Preferred Exporter for Global Buyers Miltefosine Capsules 50mg Manufacturer in India As a trusted pharmaceutical manufacturer in India, Taj Medico is committed to delivering premium-quality Miltefosine Capsules 50mg that meet stringent international quality standards. We manufacture our products under advanced production systems with a strong focus on quality assurance, consistency, safety, and regulatory compliance. Our Miltefosine Capsules 50mg (Impataj) are widely supplied across India and exported to countries in Africa, the Middle East, Latin America (LATAM), Southeast Asia, and other international markets. We support hospitals, government procurement agencies, pharmaceutical distributors, wholesalers, importers, and private label partners with reliable pharmaceutical manufacturing and export services. What Are Miltefosine Capsules 50mg? Miltefosine 50mg capsules Miltefosine Capsules 50mg belong to the class of antimicrobial medicines known as antiparasitic agents. They are prescribed for the treatment of several forms of leishmaniasis, a parasitic disease transmitted through infected sandflies. The medicine is indicated for treating: Visceral leishmaniasis (Kala-azar) Cutaneous leishmaniasis Mucosal leishmaniasis Miltefosine works by disrupting essential cellular functions required for the survival and multiplication of the parasite, ultimately leading to its destruction. Treatment should always be administered under the supervision of a […]
Zinc Acetate Tablets 50mg Manufacturer, Supplier & Exporter in India | Taj Medico Trusted Zinc Acetate Tablets 50mg Manufacturer in India Taj Medico is a trusted Zinc Acetate Tablets 50mg manufacturer, supplier, exporter, and PCD Pharma Franchise company in India, offering high-quality pharmaceutical formulations for domestic and international markets. Manufactured at our facility in Vapi, Gujarat, India, our Zinc Acetate Tablets 50mg are produced using carefully selected raw materials and stringent quality control procedures to ensure consistency, safety, and reliability. We serve pharmaceutical distributors, wholesalers, hospitals, healthcare institutions, pharmacies, and global buyers across India, Africa, the Middle East, Latin America (LATAM), and other international markets, subject to applicable regulatory requirements. Zinc acetate Tablets 50mg (Zinkotaj) What Are Zinc Acetate Tablets 50mg? Zinc Acetate Tablets 50mg contain zinc acetate, an essential mineral that supports numerous biological functions in the human body. Zinc plays an important role in normal growth, immune function, wound healing, cell division, protein synthesis, and tissue repair. Healthcare professionals prescribe Zinc Acetate Tablets 50mg to treat and prevent zinc deficiency in individuals who require zinc supplementation. When zinc levels are inadequate, supplementation may help restore normal physiological functions under the supervision of a qualified healthcare professional. Therapeutic Uses […]
Zolmitriptan Tablets 5mg Manufacturer, Supplier & Exporter in India | Taj Medico Trusted Zolmitriptan Tablets 5mg Manufacturer in India Taj Medico is a recognized manufacturer, supplier, exporter, and PCD Pharma Franchise company for Zolmitriptan Tablets 5mg in India, serving pharmaceutical distributors, hospitals, healthcare institutions, wholesalers, retailers, and international buyers. We manufacture Zolmitriptan Tablets 5mg using advanced pharmaceutical manufacturing processes and comprehensive quality control systems to deliver consistent, reliable pharmaceutical products. Our manufacturing facility in Vapi, Gujarat, India is equipped with modern production technology and follows established pharmaceutical quality standards throughout procurement, manufacturing, packaging, storage, and distribution. We are committed to supplying high-quality pharmaceutical formulations to customers across India, Africa, the Middle East, Latin America (LATAM), and other international markets, subject to applicable regulatory requirements. Zolmitriptan Tablets What Are Zolmitriptan Tablets 5mg? Zolmitriptan Tablets 5mg contain Zolmitriptan, a medicine prescribed for the acute treatment of migraine attacks with or without aura in adults. It is intended to relieve migraine symptoms after an attack has started and is not generally used to prevent future migraine episodes unless advised by a qualified healthcare professional. Migraine headaches are often associated with severe throbbing pain, nausea, vomiting, and increased sensitivity to light and sound. Zolmitriptan […]
Zolmitriptan Tablets 2.5mg Manufacturer, Supplier & Exporter in India – Trusted Pharmaceutical Partner | Taj Medico Leading Zolmitriptan Tablets 2.5mg Manufacturer in India Taj Medico is a trusted Zolmitriptan Tablets 2.5mg manufacturer, supplier, exporter, and PCD Pharma Franchise company in India, offering premium pharmaceutical products manufactured under stringent quality standards. With advanced manufacturing capabilities in Vapi, Gujarat, we supply high-quality Zolmitriptan Tablets 2.5mg across PAN India and export to healthcare organizations, pharmaceutical distributors, and wholesalers worldwide. Our commitment to quality, regulatory compliance, competitive pricing, and reliable delivery has established us as a preferred pharmaceutical manufacturing partner for domestic and international buyers. Zolmitriptan Tablets 2.5mg About Zolmitriptan Tablets 2.5mg Zolmitriptan Tablets 2.5mg belong to the triptan class of prescription medicines indicated for the acute treatment of migraine attacks with or without aura in adults. The medicine works by activating serotonin (5-HT1B/1D) receptors, helping narrow dilated blood vessels associated with migraine and reducing the release of inflammatory neuropeptides. It is intended to relieve migraine symptoms after they begin and is not indicated for migraine prevention. Therapeutic Uses Healthcare professionals prescribe Zolmitriptan Tablets 2.5mg for: Acute migraine attacks Migraine with aura Migraine without aura Relief from migraine-associated nausea Reduction of sensitivity to light […]
Anidulafungin for Injection 100mg Manufacturer in India – Trusted Global Supplier & Exporter Anidulafungin for Injection 100mg Manufacturer in India Taj Medico is a trusted Anidulafungin for Injection 100mg manufacturer in India, committed to delivering world-class injectable antifungal medicines that comply with stringent international quality standards. We manufacture and export premium-quality Anidulafungin 100mg Injection (Fungsafe) for hospitals, government institutions, pharmaceutical distributors, importers, and healthcare organizations across global markets. With advanced pharmaceutical manufacturing capabilities in Vapi, Gujarat, India, we ensure every batch is produced under strict GMP-compliant conditions using carefully selected pharmaceutical-grade raw materials. Our commitment to quality, safety, consistency, and timely delivery has established us as a preferred manufacturing and export partner throughout Africa, the Middle East, Latin America, Asia, and other international markets. What is Anidulafungin for Injection 100mg? Anidulafungin for injection 100mg Anidulafungin for Injection 100mg is a prescription antifungal medicine belonging to the echinocandin class of antifungal drugs. It is widely used for treating serious invasive fungal infections caused primarily by Candida species. The medicine inhibits fungal cell wall synthesis, resulting in rapid destruction of fungal cells while maintaining an excellent safety profile. Healthcare professionals frequently prescribe Anidulafungin for critically ill patients because of its effectiveness against […]
Ticagrelor 60mg Tablets Manufacturer & Supplier in India | Trusted Exporter and PCD Pharma Franchise Leading Ticagrelor 60mg Tablets Manufacturer in India Taj Medico is a trusted Ticagrelor 60mg Tablets Manufacturer, Supplier, Exporter, and PCD Pharma Franchise Company based in Vapi, Gujarat, India. We manufacture premium-quality cardiovascular medicines using advanced pharmaceutical technology, modern manufacturing facilities, and comprehensive quality management systems. Our commitment to product excellence, regulatory compliance, and customer satisfaction has made us a preferred pharmaceutical partner for distributors, hospitals, wholesalers, healthcare institutions, and international buyers. Our Ticagrelor 60mg Tablets are produced using carefully selected pharmaceutical ingredients and stringent quality assurance procedures. Every batch undergoes extensive quality testing to ensure consistent purity, safety, stability, and therapeutic performance before commercial release. About Ticagrelor 60mg Tablets Ticagrelor 60mg Tablets Ticagrelor 60mg Tablets are prescription-only antiplatelet medicines used to help reduce the risk of cardiovascular death, heart attack, and stroke in appropriate patients under the supervision of a healthcare professional. The medicine works by preventing platelets from sticking together, reducing the formation of harmful blood clots within blood vessels. Ticagrelor may be prescribed for selected patients who have experienced a previous heart attack or for certain patients with acute coronary syndrome (ACS), including […]
Ticagrelor Tablets 90mg Manufacturer & Supplier in India – Premium Quality Exporter and PCD Pharma Franchise Leading Ticagrelor Tablets 90mg Manufacturer in India Taj Medico is a trusted Ticagrelor Tablets 90mg manufacturer, supplier, exporter, and PCD Pharma Franchise company headquartered in Vapi, Gujarat, India. We manufacture high-quality cardiovascular medicines using advanced pharmaceutical technology, stringent quality systems, and carefully selected raw materials to ensure exceptional product quality, purity, and consistency. Our commitment to excellence has made us a preferred manufacturing partner for pharmaceutical distributors, hospitals, wholesalers, institutional buyers, and international importers. Our Ticagrelor Tablets 90mg are produced in modern manufacturing facilities that follow rigorous quality management practices. Every production batch undergoes comprehensive quality testing to ensure compliance with applicable pharmacopeial and regulatory standards before commercial distribution. About Ticagrelor Tablets 90mg Ticagrelor Tablets 90mg Ticagrelor Tablets 90mg are a prescription antiplatelet medication used to reduce the risk of serious cardiovascular events in appropriate patients. The medicine works by preventing blood platelets from sticking together, thereby reducing the formation of harmful blood clots. This mechanism helps lower the risk of heart attack, stroke, and clot-related complications when prescribed by a healthcare professional. Ticagrelor is commonly prescribed for patients who have experienced acute coronary […]
Isosorbide 5 Mononitrate BP 60mg Tablets Manufacturer & Supplier in India Trusted Manufacturer of Isosorbide 5 Mononitrate BP 60mg Tablets Taj Medico is a leading manufacturer, exporter, and supplier of Isosorbide 5 Mononitrate BP 60mg Tablets from Vapi, Gujarat, India. We manufacture high-quality pharmaceutical formulations in compliance with stringent quality standards, delivering reliable cardiovascular medicines to domestic and international markets. Our Isosorbide 5 Mononitrate BP 60mg Tablets (Tuki) are manufactured using carefully selected pharmaceutical ingredients and modern production processes to ensure consistent quality, safety, and efficacy. We serve pharmaceutical distributors, hospitals, institutional buyers, government organizations, wholesalers, and international importers by supplying premium-quality Isosorbide Mononitrate Tablets throughout PAN India, Africa, the Middle East, Latin America, Southeast Asia, and other global markets. Isosorbide 5 Mononitrate BP 60mg Tablets About Isosorbide 5 Mononitrate BP 60mg Tablets Isosorbide 5 Mononitrate BP 60mg belongs to the nitrate class of medicines used for the prevention of angina pectoris (heart-related chest pain). It works by relaxing and widening blood vessels, allowing blood to flow more easily and reducing the workload on the heart. By lowering the heart’s oxygen demand, it helps reduce the frequency of angina attacks in patients with coronary artery disease. This medicine is intended […]
Moxonidine Tablets 0.3mg Manufacturer & Supplier in India | Trusted Exporter from Vapi, Gujarat Leading Moxonidine Tablets 0.3mg Manufacturer in India Moxonidine Tablets 0.3mg are widely prescribed for the management of essential hypertension (high blood pressure). As an effective centrally acting antihypertensive medicine, moxonidine helps reduce elevated blood pressure by stimulating imidazoline receptors in the brain, thereby decreasing sympathetic nervous system activity. This mechanism supports improved cardiovascular health while helping lower the long-term risks associated with uncontrolled hypertension. We are a trusted Moxonidine Tablets 0.3mg manufacturer in India, supplying high-quality pharmaceutical products to hospitals, distributors, wholesalers, pharmacies, government institutions, and international buyers. Our manufacturing operations follow stringent quality standards, ensuring every batch meets established pharmaceutical specifications for safety, consistency, and effectiveness. Our manufacturing facility in Vapi, Gujarat serves customers across India and international markets with reliable production capacity, timely delivery, and comprehensive regulatory support. Moxonidine Tablets 0.3mg – Composition and Therapeutic Use Moxonidine Tablets 0.3mg (Bimbo) Generic Name: Moxonidine Strength: 0.3 mg Dosage Form: Oral Tablet Therapeutic Category: Antihypertensive Agent Moxonidine is prescribed for adults diagnosed with hypertension. By reducing sympathetic nerve activity, it lowers peripheral vascular resistance and assists in maintaining healthy blood pressure levels. Treatment should always be […]
Moxonidine Tablets BP 0.4mg Manufacturer in India – Trusted Supplier, Exporter and PCD Pharma Franchise by Taj Medico Moxonidine Tablets BP 0.4mg Manufacturer in India Taj Medico is a trusted Moxonidine Tablets BP 0.4mg manufacturer in India, committed to delivering high-quality pharmaceutical products that comply with international quality standards. We manufacture and supply premium-quality Moxonidine Tablets BP 0.4mg for domestic and international markets with a strong focus on product consistency, regulatory compliance, and customer satisfaction. Our modern manufacturing facilities in Vapi, Gujarat, India, operate under stringent quality management systems to ensure that every batch meets pharmacopeial specifications. We serve pharmaceutical companies, government institutions, hospitals, distributors, importers, wholesalers, and healthcare organizations across India, Africa, the Middle East, Latin America, Southeast Asia, and many other international markets. What is Moxonidine Tablets BP 0.4mg? Moxonidine Tablets BP 0.4mg (Bimbo) Moxonidine Tablets BP 0.4mg is an antihypertensive medicine used for the treatment of high blood pressure (hypertension). It acts on specific receptors within the central nervous system, reducing sympathetic nervous system activity and helping lower blood pressure effectively. Proper management of hypertension significantly reduces the risk of serious cardiovascular complications such as stroke, heart attack, kidney disease, and heart failure. Moxonidine is often prescribed […]
Mebendazole Tablets 500mg Manufacturer Supplier in India – Trusted Global Exporter of Quality Anthelmintic Medicines Leading Mebendazole Tablets 500mg Manufacturer in India Taj Medico is a leading Mebendazole Tablets 500mg manufacturer in India, committed to delivering premium-quality pharmaceutical formulations that meet international healthcare standards. We manufacture, supply, and export Mebendazole Tablets 500mg to hospitals, government institutions, pharmaceutical distributors, importers, wholesalers, and PCD partners across Africa, the Middle East, Latin America (LATAM), Asia, and other international markets. Manufactured in WHO-GMP compliant facilities, our Mebendazole Tablets are produced using high-quality Active Pharmaceutical Ingredients (API) and advanced manufacturing technology. Every batch undergoes strict quality testing to ensure purity, potency, safety, and consistent therapeutic effectiveness. Our brand Menditaj has earned recognition among healthcare professionals and pharmaceutical buyers for its quality, affordability, and reliable supply chain. Mebendazole Tablets USP 500mg What is Mebendazole Tablets 500mg? Mebendazole Tablets 500mg is an effective antiparasitic medicine used to treat a wide range of worm infections. It belongs to the benzimidazole class of anthelmintic medicines and works by preventing worms from absorbing glucose. Without glucose, the parasites lose their energy source, eventually die, and are naturally eliminated from the body. The medicine is widely prescribed for treating intestinal worm […]
Tinidazole Tablets 500mg Manufacturer in India – Trusted Global Supplier & Exporter | Taj Medico Tinidazole Tablets 500mg are widely prescribed for the treatment of bacterial and protozoal infections affecting different parts of the body. At Taj Medico, we are recognized as a leading Tinidazole Tablets 500mg Manufacturer in India, offering premium-quality pharmaceutical formulations manufactured under stringent quality standards. Our commitment to excellence, regulatory compliance, and timely global delivery has made us a preferred partner for importers, distributors, hospitals, government institutions, and pharmaceutical companies across Africa, the Middle East, Latin America, CIS countries, and other international markets. Our advanced manufacturing capabilities, experienced pharmaceutical professionals, and strict quality assurance systems ensure that every batch of (Tinidataj) Tinidazole Tablets 500mg delivers consistent efficacy, safety, and reliability. Leading Tinidazole Tablets 500mg Manufacturer in India Tinidazole Tablets USP 500mg Taj Medico manufactures Tinidazole Tablets 500mg using high-quality active pharmaceutical ingredients sourced from certified vendors. Every production stage is carefully monitored to ensure compliance with WHO-GMP manufacturing guidelines and international pharmaceutical standards. We specialize in manufacturing: Tinidazole Tablets 500mg Tinidazole Tablets for Government Supply Tinidazole Tablets for Institutional Supply Tinidazole Tablets for Export Markets Private Label Tinidazole Tablets Third Party Manufacturing Contract Manufacturing PCD Pharma […]
Zinc Oxide Cream 32% Manufacturer in India | Leading Supplier, Exporter & PCD Pharma Franchise – Taj Medico Zinc Oxide Cream 32% is a widely trusted topical skin protectant formulated to provide an effective protective barrier against moisture, irritation, and external contaminants. At Taj Medico, we manufacture premium-quality Zinc Oxide Cream 32% using pharmaceutical-grade raw materials under stringent quality standards. As a leading Zinc Oxide Cream Manufacturer in India, we supply high-quality formulations to pharmaceutical companies, hospitals, distributors, wholesalers, government institutions, and international buyers across Africa, the Middle East, Latin America (LATAM), Asia, and other global markets. Our manufacturing facility in Vapi, Gujarat, India follows advanced manufacturing practices with strict quality assurance protocols to ensure consistency, purity, stability, and safety in every batch. We provide complete pharmaceutical manufacturing solutions including third-party manufacturing, contract manufacturing, private labeling, bulk supply, and PCD Pharma Franchise services. Zinc oxide cream Leading Zinc Oxide Cream 32% Manufacturer in India We have established ourselves as one of the trusted manufacturers of Zinc Oxide Cream 32% by consistently delivering products that comply with international pharmaceutical quality standards. Our experienced production team utilizes modern manufacturing technology and carefully selected ingredients to produce creams that meet the expectations of […]
Atazanavir 300mg and Ritonavir 100mg Tablets (Aifotaj-R) Manufacturer, Supplier & Exporter in India – Taj Medico Trusted Manufacturer of Atazanavir 300mg and Ritonavir 100mg Tablets in India Taj Medico is engaged in the manufacturing, supplying, exporting, and contract manufacturing of Atazanavir 300mg and Ritonavir 100mg Tablets under the brand Aifotaj-R. We support pharmaceutical distributors, healthcare institutions, government procurement agencies, and international buyers with high-quality pharmaceutical formulations manufactured under stringent quality systems. Our manufacturing operations are designed to meet recognized pharmaceutical quality standards while offering reliable supply capabilities for domestic and international markets. Through our commitment to product quality, regulatory compliance, and dependable customer support, we have established long-term partnerships with pharmaceutical companies across India and overseas. Atazanavir 300mg and Ritonavir 100mg Tablets About Atazanavir 300mg and Ritonavir 100mg Tablets Atazanavir is an antiretroviral medicine classified as a protease inhibitor. When administered with a low dose of Ritonavir, the combination helps maintain therapeutic drug concentrations by slowing the metabolism of Atazanavir. This combination is prescribed as part of combination antiretroviral therapy (ART) for the treatment of HIV-1 infection in adults and eligible patients, in accordance with current medical guidelines and under the supervision of qualified healthcare professionals. Treatment decisions, dosage, and […]
Lopinavir 100mg and Ritonavir 25mg Tablets Manufacturer Supplier in India – Trusted Global Exporter Lopinavir 100mg and Ritonavir 25mg Tablets are widely prescribed antiretroviral medicines used in the management of Human Immunodeficiency Virus (HIV) infection. At Taj Medico, we are committed to manufacturing and exporting premium-quality pharmaceutical formulations that comply with international quality standards. As a leading Lopinavir 100mg and Ritonavir 25mg Tablets Manufacturer in India, we supply high-quality formulations to hospitals, government organizations, pharmaceutical distributors, NGOs, and healthcare institutions across Africa, the Middle East, Latin America, Asia, and many other international markets. Our advanced manufacturing facilities in Vapi, Gujarat, India, operate under stringent quality management systems to ensure every batch meets global regulatory expectations. We combine scientific expertise, modern manufacturing technology, and reliable logistics to deliver pharmaceutical products that healthcare providers trust worldwide. Leading Lopinavir 100mg and Ritonavir 25mg Tablets Manufacturer in India Lopinavir 100mg and Ritonavir 25mg Tablets We have established ourselves as one of India’s trusted manufacturers and exporters of Lopinavir 100mg and Ritonavir 25mg Tablets. Our manufacturing process follows current Good Manufacturing Practices (cGMP), ensuring product consistency, safety, efficacy, and purity. Healthcare organizations choose us because we provide: WHO-GMP quality manufacturing International export standards Bulk manufacturing […]
Oxybutynin 2.5mg Tablets Manufacturer and Supplier in India – Trusted Global Exporter of Oxybutynin 2.5mg Tablets Oxybutynin 2.5mg Tablets are widely prescribed for the management of overactive bladder (OAB) and urinary incontinence. As a leading Oxybutynin 2.5mg Tablets Manufacturer in India, we deliver high-quality pharmaceutical formulations manufactured in WHO-GMP compliant facilities and supplied to healthcare organizations, importers, distributors, hospitals, government institutions, and pharmaceutical companies across the globe. Our manufacturing processes comply with stringent international quality standards, ensuring consistent efficacy, purity, safety, and stability. We are committed to providing affordable, export-quality Oxybutynin tablets with complete regulatory support for international markets. Oxybutynin 2.5mg Tablets What are Oxybutynin 2.5mg Tablets? Oxybutynin 2.5mg Tablets contain Oxybutynin Hydrochloride, an anticholinergic and antispasmodic medicine that relaxes the bladder muscles. It reduces involuntary bladder contractions, helping patients control urinary urgency, frequent urination, and urinary leakage. Oxybutynin works by blocking muscarinic receptors responsible for bladder muscle contractions, thereby increasing bladder capacity and decreasing episodes of urgency and incontinence. Medical Uses of Oxybutynin 2.5mg Tablets Oxybutynin 2.5mg Tablets are commonly prescribed for: Overactive bladder (OAB) Urinary urgency Frequent urination Urge urinary incontinence Neurogenic bladder Bladder muscle spasms Pediatric bladder dysfunction (under physician supervision) Post-surgical bladder spasms The medicine significantly […]
Leading Oxybutynin 5mg Tablets Manufacturer & Supplier in India Premium Oxybutynin 5mg Tablets Manufacturer, Supplier, Exporter & PCD Pharma Franchise in India Taj Medico is a trusted name in the Indian pharmaceutical industry, recognized as a leading Oxybutynin 5mg Tablets Manufacturer, Supplier, Exporter, and PCD Pharma Franchise Company in India. With a strong commitment to quality, innovation, and customer satisfaction, we manufacture premium-quality Oxybutynin 5mg Tablets using advanced pharmaceutical technology and stringent quality control processes. Our manufacturing facilities are equipped with modern machinery and operated by experienced pharmaceutical professionals, enabling us to deliver products that meet the highest industry standards. We supply our products across PAN India and export to international markets including Africa, the Middle East, Latin America (LATAM), and other global destinations. What are Oxybutynin 5mg Tablets? Oxybutynin 5mg Tablets Oxybutynin 5mg Tablets contain Oxybutynin Hydrochloride 5 mg, an antimuscarinic medication widely prescribed for the treatment of overactive bladder (OAB). The medicine works by relaxing the bladder muscles, reducing involuntary contractions, and increasing bladder capacity. Doctors commonly prescribe Oxybutynin to help manage: Overactive bladder (OAB) Urinary urgency Frequent urination Urge urinary incontinence Bladder muscle spasms Neurogenic bladder disorders By reducing bladder contractions, Oxybutynin helps patients regain better control […]
Bedaquiline Tablets 100mg Manufacturer, Supplier & Exporter in India Leading Bedaquiline Tablets 100mg Manufacturer in India We are a trusted and globally recognized Bedaquiline Tablets 100mg Manufacturer, Supplier, and Exporter in India, committed to delivering premium-quality pharmaceutical formulations for the treatment of multidrug-resistant tuberculosis (MDR-TB). Manufactured under strict compliance standards, our products are designed to ensure high purity, accurate dosage, and consistent therapeutic performance. As a reputed pharmaceutical organization, we ensure that every batch of Bedaquiline Tablets 100mg is developed using advanced formulation technology and validated manufacturing processes that meet WHO-GMP and ISO-certified standards. About Taj Medico – Trusted Pharma Manufacturer from Vapi, Gujarat Bedaquiline Tablets 100mg Taj Medico is a well-established name in the pharmaceutical industry, known for its excellence in manufacturing, exporting, and supplying Bedaquiline Tablets 100mg across India and global markets. Based in Vapi, Gujarat, we operate with a strong infrastructure that supports large-scale production and efficient distribution. Our company is dedicated to delivering high-quality anti-tuberculosis formulations that meet international healthcare requirements, making us a preferred partner for distributors, hospitals, and global healthcare suppliers. Bedaquiline Tablets 100mg – Composition and Therapeutic Use Bedaquiline is a specialized anti-tuberculosis medication used in combination therapy for pulmonary tuberculosis caused by […]
Glucagon for Injection 1mg Manufacturer Supplier in India – Leading Gluvio Exporter, Supplier & PCD Pharma Franchise Partner Glucagon for Injection 1mg Manufacturer in India – Trusted Quality from Taj Medico Glucagon for Injection 1mg is a critical emergency medication widely used for the treatment of severe hypoglycemia in patients with diabetes. When blood sugar levels fall dangerously low and the patient becomes unconscious or unable to consume glucose orally, Glucagon Injection acts rapidly to increase blood sugar levels and restore consciousness. Due to its life-saving role in emergency diabetes management, the global demand for high-quality Glucagon for Injection 1mg continues to rise across hospitals, clinics, emergency care centers, healthcare institutions, and pharmaceutical distributors. Taj Medico, the well-known Glucagon for Injection 1mg Manufacturer in India, provides assurance of exceptional quality, purity, efficacy, and safety. Manufactured under stringent pharmaceutical quality standards, our Glucagon Injection is trusted by healthcare professionals and pharmaceutical buyers worldwide. As a leading Glucagon for Injection 1mg Supplier, Manufacturer, Exporter, and PCD Pharma Franchise Provider, we serve clients across India, Africa, the Middle East, Latin America, CIS countries, and various emerging pharmaceutical markets. What is Glucagon for Injection 1mg? Glucagon for Injection 1mg Glucagon for Injection 1mg (Gluvio) […]
Ciprofloxacin Tablets 500mg Manufacturer Supplier and PCD Pharma Franchise in India – Leading Exporter and Trusted Pharmaceutical Partner Ciprofloxacin Tablets 500mg Manufacturer in India – Premium Quality Pharmaceutical Solutions Ciprofloxacin Tablets 500mg are among the most widely prescribed antibacterial medicines used for the treatment of various bacterial infections. As the demand for high-quality pharmaceutical products continues to increase across domestic and international markets, healthcare providers, distributors, wholesalers, hospitals, and importers are actively seeking reliable manufacturers capable of delivering consistent quality and regulatory compliance. Taj Medico has established itself as a leading Ciprofloxacin Tablets 500mg Manufacturer in India, offering superior-quality formulations manufactured under stringent quality control systems. With advanced manufacturing facilities, experienced pharmaceutical professionals, and global export capabilities, we have become a preferred partner for buyers seeking dependable pharmaceutical products. Our commitment to quality, affordability, and timely delivery has enabled us to serve clients throughout India and across international markets. We manufacture Ciprofloxacin Tablets 500mg according to internationally accepted pharmaceutical standards, ensuring efficacy, safety, and consistency in every batch. What is Ciprofloxacin Tablets 500mg? Ciprofloxacin Tablets USP 500mg Ciprofloxacin 500mg belongs to the fluoroquinolone class of antibiotics and is used to treat a broad range of bacterial infections. It works by […]
Sildenafil 100mg Tablets Manufacturer Supplier in India – Leading Exporter, Supplier and PCD Pharma Franchise Company 🏭 Sildenafil 100mg Tablets Manufacturer in India Taj Medico is a renowned Sildenafil 100mg Tablets Manufacturer in India, offering premium-quality pharmaceutical formulations that meet stringent international quality standards. We are recognized as a trusted Manufacturer, Supplier, Exporter, and PCD Pharma Franchise Company delivering high-quality Sildenafil 100mg Tablets (Neogra) to pharmaceutical distributors, wholesalers, hospitals, pharmacies, healthcare institutions, and importers across global markets. Manufactured at our advanced pharmaceutical facility in Vapi, Gujarat, Sildenafil 100mg Tablets are produced using high-quality active pharmaceutical ingredients and modern manufacturing technologies. Every batch undergoes rigorous quality testing to ensure safety, efficacy, purity, and consistency. As a leading Sildenafil 100mg Tablets Manufacturer Supplier and Exporter in India, Taj Medico serves domestic and international pharmaceutical markets including Africa, the Middle East, Latin America, Southeast Asia, and CIS countries. 💊 What Are Sildenafil 100mg Tablets? Sildenafil 100mg Tablets Sildenafil 100mg Tablets are prescription medicines primarily indicated for the treatment of Erectile Dysfunction (ED) in adult men. Erectile dysfunction is a condition characterized by the inability to achieve or maintain an erection sufficient for satisfactory sexual activity. Sildenafil belongs to a group of medicines known […]
Sofosbuvir 400mg and Velpatasvir 100mg Tablets Manufacturer Supplier in India Sofosbuvir 400mg and Velpatasvir 100mg Tablets Manufacturer in India Taj Medico, the well-known Sofosbuvir 400mg and Velpatasvir 100mg Tablets Manufacturer in India, is committed to delivering superior pharmaceutical products that meet international quality standards. We are recognized as a trusted Manufacturer, Supplier, Exporter, and PCD Pharma Franchise Company offering premium-quality antiviral medicines to healthcare institutions, distributors, wholesalers, pharmacies, hospitals, and pharmaceutical importers worldwide. Our flagship antiviral formulation, Sofosbuvir 400mg and Velpatasvir 100mg Tablets, marketed under the brand Hepcitaj-V, is widely used in the treatment of chronic Hepatitis C Virus (HCV) infection. Manufactured in advanced pharmaceutical facilities located in Vapi, Gujarat, India, these tablets are produced using stringent quality control measures to ensure safety, efficacy, stability, and therapeutic reliability. As a leading Sofosbuvir and Velpatasvir Tablets Manufacturer from Mumbai and Vapi, we serve domestic and international pharmaceutical markets across Africa, the Middle East, Latin America, Asia, and other emerging healthcare regions. What Are Sofosbuvir 400mg and Velpatasvir 100mg Tablets? Sofosbuvir 400mg and Velpatasvir 100mg Tablets Sofosbuvir 400mg and Velpatasvir 100mg Tablets are a fixed-dose combination antiviral medication designed for the treatment of chronic Hepatitis C infection. This powerful combination contains: Sofosbuvir […]
Sildenafil 50mg Tablets Manufacturer Supplier and Exporter in India Leading Sildenafil 50mg Tablets Manufacturer in India Taj Medico is a trusted and reputed Sildenafil 50mg Tablets Manufacturer, Supplier, Exporter, and PCD Pharma Franchise Company in India, offering premium-quality pharmaceutical products to domestic and international markets. With advanced manufacturing facilities, stringent quality control systems, and a commitment to excellence, we have established ourselves as one of the leading suppliers of Sildenafil 50mg Tablets (Neogra) across India, Africa, the Middle East, and Latin America. Our Sildenafil 50mg Tablets are manufactured using high-purity Sildenafil Citrate and comply with international quality standards. We focus on delivering safe, effective, and affordable formulations that meet the evolving needs of healthcare professionals, distributors, importers, and pharmaceutical businesses worldwide. Sildenafil 50mg Tablets What Are Sildenafil 50mg Tablets? Sildenafil 50mg Tablets are prescription medications primarily used for the treatment of erectile dysfunction (ED) in men. The medicine works by increasing blood flow to the penile region, helping men achieve and maintain an erection during sexual stimulation. Sildenafil belongs to a class of medicines known as Phosphodiesterase Type 5 (PDE5) Inhibitors. It relaxes the blood vessels and improves circulation, making it one of the most prescribed treatments for erectile dysfunction […]
Ciprofloxacin Tablets 250mg Manufacturer in India Ciprofloxacin Tablets 250mg Manufacturer and Exporter in India India has established itself as one of the world’s most trusted pharmaceutical manufacturing destinations, supplying high-quality medicines to healthcare providers, distributors, hospitals, government organizations, and pharmaceutical companies across international markets. Among the essential antibacterial medicines produced in India, Ciprofloxacin Tablets 250mg remain an important therapeutic option for treating a wide range of bacterial infections. At Taj Medico, we manufacture Ciprofloxacin Tablets 250mg using advanced pharmaceutical manufacturing processes and rigorous quality control systems. Our objective is to deliver consistent product quality, regulatory compliance, and dependable supply solutions to domestic and international customers. We serve clients across India as well as export markets in Africa, the Middle East, Latin America (LATAM), Southeast Asia, and other global regions. Ciprofloxacin Tablets 250mg What Are Ciprofloxacin Tablets 250mg? Ciprofloxacin is a fluoroquinolone antibiotic that is prescribed for the treatment of various bacterial infections. It works by inhibiting bacterial DNA replication, preventing bacteria from multiplying and allowing the body’s immune system to eliminate the infection effectively. Ciprofloxacin Tablets 250mg are intended for use only under the supervision of a qualified healthcare professional. Since antibiotics are effective only against bacterial infections, they should […]
Sulfinpyrazone Tablets 200mg Manufacturer, Supplier & Exporter in India Introduction to Sulfinpyrazone Tablets 200mg Production Sulfinpyrazone Tablets 200mg are widely used in the management of chronic gout and uric acid–related disorders. As a uricosuric medication, Sulfinpyrazone supports the reduction of serum uric acid levels by enhancing renal excretion. This makes it an important therapeutic option in long-term care of patients suffering from Gout. We operate as a specialized manufacturer, exporter, and supplier of Sulfinpyrazone Tablets 200mg in India, delivering pharmaceutical-grade formulations designed for domestic and international healthcare markets. Our focus remains on precision manufacturing, regulatory compliance, and consistent product quality across every production batch. Sulfinpyrazone Tablets 200mg Composition and Therapeutic Role Sulfinpyrazone Tablet 200mg Sulfinpyrazone is classified as a uricosuric agent that assists in lowering uric acid levels in the bloodstream. The tablets are formulated to ensure controlled release and optimal absorption, supporting long-term urate management. Therapeutic Functions Reduction of elevated uric acid levels Prevention of urate crystal formation in joints Long-term suppression of recurrent gout attacks Supportive therapy for chronic hyperuricemia The medication does not provide immediate relief from acute gout pain but is highly effective in preventing future episodes when used consistently under medical supervision. Mechanism of Action […]
Amlodipine Besylate 10mg Tablets PAN India Manufacturer Introduction to Amlodipine Besylate 10mg Tablets Manufacturing Excellence We are a leading manufacturer, exporter, and supplier of Amlodipine Besylate 10mg Tablets in India, delivering high-quality pharmaceutical formulations that meet stringent global standards. Our products are developed under advanced manufacturing practices, ensuring safety, efficacy, and consistency across every batch. Amlodipine Besylate 10mg Tablets are widely prescribed as a calcium channel blocker used in the management of hypertension and angina, helping patients achieve better cardiovascular health and improved quality of life. We are committed to providing reliable pharmaceutical solutions that support healthcare providers and distributors across India, Africa, the Middle East, and Latin America. Our focus remains on delivering premium-grade formulations that meet international regulatory expectations. What is Amlodipine Besylate 10mg Tablets? Amlodipine besylate Tablets Amlodipine Besylate is a long-acting calcium channel blocker used primarily in the treatment of high blood pressure (hypertension) and chest pain (angina). It works by relaxing and widening blood vessels, allowing smoother blood flow and reducing the workload on the heart. By improving blood circulation, Amlodipine helps lower blood pressure levels and reduces the risk of heart-related complications. It is widely used due to its once-daily dosage convenience and long-lasting […]
Cetirizine 10mg Tablets High-Quality Antihistamine Manufacturing & Supply Introduction of Cetirizine 10mg Tablets We are engaged in the manufacturing, supplying, and export of Cetirizine 10mg tablets formulated to provide effective relief from a wide range of allergic conditions. Cetirizine is a second-generation antihistamine widely used for managing symptoms associated with seasonal allergies, perennial allergic rhinitis, and chronic. Our Cetirizine tablets are developed under strict pharmaceutical quality systems to ensure safety, stability, and therapeutic effectiveness. Each tablet is carefully formulated to deliver consistent dosage accuracy and reliable patient outcomes. Cetirizine works by selectively blocking peripheral H1 histamine receptors, thereby reducing the body’s allergic response without causing significant sedation in most users. Therapeutic Uses of Cetirizine Tablets Cetirizine 10mg Tablets Cetirizine 10mg tablets are widely recommended for the management of various allergic conditions. We ensure that our formulation is suitable for both clinical and general healthcare use. Common Indications We provide Cetirizine tablets for relief from: Allergic rhinitis (seasonal and perennial) Sneezing and nasal congestion Runny nose due to allergies Itchy or watery eyes Skin allergies and urticaria (hives) Allergic reactions caused by insect bites Mild dermatological allergic responses These tablets are widely used in both acute symptom management and long-term allergy […]
Topiramate 25mg Tablets: Uses, Manufacturing Standards, and Pharmaceutical Overview Introduction to Topiramate 25mg Tablets Topiramate 25mg tablets are a widely used antiepileptic medication prescribed for the management of neurological disorders such as epilepsy and the prevention of migraine headaches. The active pharmaceutical ingredient, Topiramate, acts on the central nervous system by stabilizing neuronal activity and reducing excessive electrical discharges in the brain. This medication is commonly prescribed as part of long-term therapy and is often initiated at low doses such as 25mg to minimize side effects and allow gradual dose adjustment based on patient response. Therapeutic Uses of Topiramate Topiramate 25mg Tablets Topiramate is clinically indicated for several neurological and related conditions, including: 1. Epilepsy Management Topiramate is used either as monotherapy or adjunct therapy for controlling seizures. It is effective against: Partial-onset seizures Generalized tonic-clonic seizures Seizures associated with Lennox-Gastaut syndrome It works by modulating sodium channels, enhancing GABA activity, and inhibiting excitatory neurotransmission. 2. Migraine Prevention Topiramate is widely prescribed for reducing the frequency of migraine attacks. It does not treat acute headaches but helps in long-term prevention by stabilizing neural excitability. 3. Off-label Applications In some cases, under strict medical supervision, Topiramate may be used for: Weight […]
Clarithromycin 250mg Tablets Manufacturer, Exporter & Supplier in India Leading Manufacturer of Clarithromycin 250mg Tablets for Global Healthcare Needs We are a trusted and established manufacturer, exporter, and supplier of Clarithromycin 250mg Tablets in India, delivering high-quality pharmaceutical formulations that meet stringent international standards. Our manufacturing facility is designed to ensure consistent production of safe, effective, and pure macrolide antibiotic tablets used widely in the treatment of bacterial infections. Clarithromycin is a well-recognized antibiotic belonging to the macrolide class, prescribed for a wide range of bacterial infections affecting the respiratory tract, skin, throat, ears, sinuses, and lungs. With a strong commitment to pharmaceutical excellence, we ensure every batch of Clarithromycin tablets delivers optimal therapeutic performance. About Clarithromycin 250mg Tablets Clarithromycin 250mg Tablets Clarithromycin 250mg Tablets are an effective antibiotic medication that works by inhibiting bacterial protein synthesis. By blocking the ribosomal function of bacteria, it prevents their growth and multiplication, thereby controlling and eliminating infection. This makes it highly effective in treating conditions such as: Tonsillitis and pharyngitis Sinusitis and ear infections Bronchitis and community-acquired pneumonia Skin and soft tissue infections Certain atypical bacterial infections in immunocompromised patients The drug is known for its broad-spectrum activity against Gram-positive and some […]
Ibuprofen Tablets 400mg Manufacturer, Supplier & Exporter in India Ibuprofen Tablets 400mg Manufacturer in India We are a leading Ibuprofen Tablets 400mg manufacturer, supplier, and exporter in India, committed to delivering high-quality pharmaceutical products that meet international standards. Ibuprofen Tablets 400mg, marketed under the brand name Ibrutaj, are widely used for pain relief, fever reduction, and management of inflammatory conditions. With advanced manufacturing facilities, stringent quality control processes, and global distribution capabilities, we ensure that every batch of Ibuprofen Tablets 400mg maintains the highest standards of safety, efficacy, and purity. As a trusted pharmaceutical company based in Vapi, Gujarat, India, we serve customers across India, Africa, the Middle East, Latin America, and various international markets. Our expertise in pharmaceutical manufacturing enables us to provide reliable products for healthcare professionals, distributors, hospitals, pharmacies, and PCD pharma franchise partners. What is Ibuprofen Tablets 400mg? Ibuprofen 400mg Tablets Ibuprofen Tablets 400mg is a non-steroidal anti-inflammatory drug (NSAID) that helps reduce pain, inflammation, and fever. It works by blocking the production of prostaglandins, substances responsible for pain and inflammation in the body. Ibuprofen is commonly prescribed for: Headache Migraine Fever Toothache Menstrual pain Muscle pain Joint pain Arthritis Back pain Cold and flu symptoms […]
Biperiden Tablets 4mg Manufacturer, Supplier and Exporter in India Biperiden Tablets 4mg Manufacturer and Supplier in India Taj Medico is a trusted name in the pharmaceutical industry, recognized as a leading manufacturer, supplier, and exporter of Biperiden Tablets 4mg (Inpaxid) in India. With a strong commitment to quality, innovation, and regulatory compliance, we provide premium pharmaceutical formulations that meet international standards. Our Biperiden Tablets are manufactured in advanced facilities using high-quality active pharmaceutical ingredients to ensure efficacy, safety, and consistency. We serve pharmaceutical distributors, healthcare institutions, wholesalers, government agencies, and PCD pharma franchise partners across India and international markets including Africa, the Middle East, and Latin America. Our dedication to excellence has positioned us among the preferred Biperiden Tablets manufacturers in India. Biperiden Hydrochloride Tablets 4mg What is Biperiden Tablets 4mg? Biperiden Tablets 4mg is an anticholinergic medication primarily prescribed for the treatment of Parkinson’s disease and drug-induced extrapyramidal symptoms. The medicine helps manage symptoms such as muscle rigidity, tremors, stiffness, involuntary movements, and difficulties associated with motor control. By restoring the balance of neurotransmitters in the brain, Biperiden supports improved muscle coordination and helps patients maintain better physical mobility. The medication is widely prescribed by healthcare professionals for managing […]
Misoprostol 200mcg Tablets Manufacturer in India Misoprostol 200mcg Tablets Manufacturer and Supplier in India India has emerged as one of the world’s most reliable pharmaceutical manufacturing hubs, offering high-quality formulations, advanced production facilities, and cost-effective healthcare solutions. Among the leading pharmaceutical products supplied globally, Misoprostol 200mcg Tablets continue to play an important role in modern healthcare applications where prescribed by qualified medical professionals. As a trusted Misoprostol 200mcg Tablets Manufacturer in India, Taj Medico focuses on quality-driven manufacturing, regulatory compliance, dependable supply chains, and international distribution support. Our commitment to excellence has enabled us to serve pharmaceutical distributors, wholesalers, healthcare organizations, importers, and business partners across multiple international markets. Misoprostol 200mcg Tablets Exporter from India Misoprostol Tablets 200mcg The demand for pharmaceutical products manufactured in India continues to grow across Africa, Latin America, Asia, and the Middle East. Taj Medico has established a strong reputation as a reliable exporter of Misoprostol 200mcg Tablets by maintaining strict quality standards and ensuring consistency throughout the manufacturing process. Our export operations are supported by: Modern manufacturing infrastructure Stringent quality assurance systems Timely order fulfillment International packaging standards Reliable logistics management Professional documentation support Global distribution capabilities By maintaining these standards, we help international […]
Chlorthalidone 25mg Tablets Manufacturer in India Leading Chlorthalidone 25mg Tablets Manufacturer in India Taj Medico is a renowned Chlorthalidone 25mg Tablets Manufacturer in India, committed to delivering high-quality pharmaceutical formulations that meet stringent international standards. We specialize in manufacturing, exporting, and supplying Chlorthalidone Tablets to healthcare institutions, distributors, importers, wholesalers, government organizations, and pharmaceutical partners across India, Africa, the Middle East, Latin America, and other global markets. Our manufacturing facilities are equipped with advanced technology, modern quality control systems, and highly skilled pharmaceutical professionals dedicated to ensuring product consistency, safety, efficacy, and compliance. We maintain strict adherence to international pharmaceutical manufacturing practices to ensure superior product quality. What is Chlorthalidone 25mg Tablets? Chlorthalidone 25mg Tablets Chlorthalidone 25mg Tablets is a thiazide-like diuretic medication widely prescribed for the treatment of: Hypertension (High Blood Pressure) Edema associated with Heart Failure Kidney Disorders Liver Disorders Fluid Retention Conditions Cardiovascular Risk Reduction Chlorthalidone works by increasing urine production, helping the body eliminate excess salt and water. This mechanism helps lower blood pressure and reduces swelling caused by fluid accumulation. Benefits of Chlorthalidone 25mg Tablets Chlorthalidone is recognized worldwide for its effectiveness in managing hypertension and edema. Major Benefits Include: Effective blood pressure control Long-lasting […]
Riociguat 0.5mg Tablets Manufacturer in India Riociguat 0.5mg Tablets Manufacturer and Exporter in India Taj Medico is a leading Riociguat 0.5mg Tablets Manufacturer in India, delivering world-class pharmaceutical products that meet stringent international quality standards. With a strong presence in regulated and semi-regulated markets, we have established ourselves as a trusted partner for healthcare institutions, pharmaceutical distributors, government tenders, and PCD pharma franchise companies across the globe. Our Riociguat 0.5mg Tablets are manufactured in state-of-the-art facilities that comply with WHO-GMP standards and international regulatory requirements. We are committed to ensuring product quality, efficacy, safety, and consistency in every batch supplied to domestic and international markets. What is Riociguat 0.5mg Tablet? Riociguat 0.5mg Tablets Riociguat 0.5mg Tablet is a prescription medication primarily used in the treatment of: Chronic Thromboembolic Pulmonary Hypertension (CTEPH) Pulmonary Arterial Hypertension (PAH) The medicine works by stimulating soluble guanylate cyclase (sGC), helping blood vessels in the lungs relax and widen. This improves blood flow and reduces pressure within pulmonary arteries, allowing patients to improve exercise capacity and overall quality of life. Riociguat is recognized globally as an effective therapeutic option for managing pulmonary hypertension and related cardiovascular conditions. Leading Riociguat Tablets Manufacturer in India As a reputed […]
Riociguat 1mg Tablets Manufacturer in India Riociguat 1mg Tablets Manufacturer, Supplier and Exporter in India We are recognized as one of the leading Riociguat 1mg Tablets Manufacturers in India, offering high-quality pharmaceutical formulations that meet international quality standards. Our Riociguat Tablets are manufactured in WHO-GMP compliant facilities using premium-grade active pharmaceutical ingredients and advanced production technologies to ensure safety, efficacy, and consistency. Riociguat is an important prescription medication used for the treatment of pulmonary arterial hypertension (PAH) and chronic thromboembolic pulmonary hypertension (CTEPH). As a trusted pharmaceutical company, we supply and export Riociguat 1mg Tablets to regulated and semi-regulated markets across Africa, the Middle East, Latin America, Asia, and other global destinations. Our commitment to quality manufacturing, regulatory compliance, competitive pricing, and timely delivery has made us a preferred partner for distributors, healthcare institutions, importers, wholesalers, government tenders, and pharmaceutical companies worldwide. What is Riociguat 1mg Tablet? Riociguat 1mg Tablets Riociguat is a soluble guanylate cyclase (sGC) stimulator used for treating specific forms of pulmonary hypertension. It works by relaxing blood vessels in the lungs, improving blood flow, reducing pressure in pulmonary arteries, and enhancing exercise capacity in patients suffering from pulmonary hypertension. Riociguat 1mg Tablets are prescribed for: Pulmonary […]
Riociguat 2mg Tablets Manufacturer in India Riociguat 2mg Tablets Manufacturer and Supplier in India India has emerged as one of the world’s leading pharmaceutical manufacturing hubs, supplying high-quality medicines to domestic and international healthcare markets. Among specialized cardiovascular and pulmonary care medications, Riociguat 2mg Tablets play a critical role in the treatment of Pulmonary Arterial Hypertension (PAH) and Chronic Thromboembolic Pulmonary Hypertension (CTEPH). As a trusted Riociguat 2mg Tablets manufacturer in India, we focus on delivering pharmaceutical products that meet stringent quality standards, ensuring safety, efficacy, and consistency in every batch. Our manufacturing operations are designed to comply with international pharmaceutical guidelines, enabling healthcare providers, distributors, hospitals, pharmacies, and pharmaceutical companies to access dependable Riociguat tablet supplies across India and global markets. What Are Riociguat 2mg Tablets? Riociguat Tablets 2mg Riociguat is an advanced prescription medication used in the management of pulmonary hypertension disorders. It works by stimulating soluble guanylate cyclase (sGC), helping blood vessels in the lungs relax and widen. This action improves blood circulation, reduces pressure within pulmonary arteries, and enhances exercise capacity in patients suffering from pulmonary vascular diseases. Riociguat tablets are commonly prescribed for: Pulmonary Arterial Hypertension (PAH) Chronic Thromboembolic Pulmonary Hypertension (CTEPH) Long-term pulmonary hypertension […]
Mexiletine Hydrochloride Capsules 200mg Manufacturer in India Leading Mexiletine Hydrochloride Capsules 200mg Manufacturer in India India has emerged as a global hub for high-quality pharmaceutical manufacturing, serving healthcare markets across Asia, Africa, the Middle East, Latin America, and Europe. Among specialized cardiac medications, Mexiletine Hydrochloride Capsules 200mg remain an important treatment option for managing serious ventricular arrhythmias and other cardiac rhythm disorders. As a trusted Mexiletine Hydrochloride Capsules 200mg Manufacturer in India, we are committed to delivering pharmaceutical products that meet international quality standards, regulatory requirements, and global healthcare demands. Our advanced manufacturing facilities, stringent quality control systems, and dedicated supply chain management enable us to provide reliable pharmaceutical solutions to distributors, hospitals, wholesalers, healthcare institutions, and pharmaceutical importers worldwide. Mexiletine Capsules 200mg What Are Mexiletine Hydrochloride Capsules 200mg? Mexiletine Hydrochloride Capsules 200mg is an oral antiarrhythmic medication used primarily in the treatment and management of life-threatening ventricular arrhythmias. The medication belongs to Class IB antiarrhythmic agents and functions by regulating abnormal electrical activity within the heart. By blocking sodium channels responsible for abnormal cardiac conduction, Healthcare professionals may also prescribe the medication for selected off-label applications under appropriate medical supervision. Therapeutic Uses of Mexiletine Hydrochloride Capsules 200mg Mexiletine Hydrochloride […]
Modafinil Tablets 100mg Manufacturer Supplier in India – Trusted Exporter & Global Pharma Partner Modafinil Tablets 100mg Manufacturer in India We are recognized among the leading manufacturers and exporters of Modafinil Tablets 100mg in India, delivering premium-quality pharmaceutical formulations for domestic and international markets. Our manufacturing facilities follow strict quality standards, advanced production practices, and international regulatory guidelines to ensure safe, effective, and high-purity medicines for global healthcare requirements. As a trusted pharmaceutical company based in Vapi, Gujarat, India, we specialize in manufacturing and supplying high-quality Modafinil Tablets 100mg for hospitals, pharmacies, distributors, importers, wholesalers, government tenders, and PCD pharma franchise businesses across worldwide markets. Our commitment to pharmaceutical excellence, affordable pricing, timely delivery, and export-quality packaging has positioned us among the preferred exporters of Modafinil Tablets in Africa, the Middle East, LATAM countries, Southeast Asia, and multiple regulated and semi-regulated markets. What is Modafinil Tablets 100mg? Modafinil Tablets 100mg Modafinil Tablets 100mg is a prescription medication commonly used for promoting wakefulness and improving alertness in individuals experiencing excessive daytime sleepiness. It is widely prescribed for conditions associated with sleep disorders and fatigue-related health conditions. This medicine helps improve concentration, focus, mental performance, and wakefulness by influencing specific neurotransmitters in […]
Modafinil Tablets USP 200mg Manufacturer, Supplier & Exporter in India Modafinil Tablets USP 200mg Manufacturer in India We are a reputed and globally recognized manufacturer, exporter, and supplier of Modafinil Tablets USP 200mg in India, delivering premium pharmaceutical formulations designed to meet international quality standards. Our manufacturing excellence is driven by advanced production infrastructure, strict quality control systems, and compliance with WHO-GMP guidelines. As a trusted pharmaceutical organization, Taj Medico is committed to providing high-purity Modafinil Tablets USP 200mg that support effective management of sleep-related disorders and excessive daytime sleepiness. Our products are widely supplied across Africa, the Middle East, Latin America (LTAM), and multiple regulated and semi-regulated markets, ensuring global healthcare accessibility. What is Modafinil Tablets USP 200mg? Modafinil Tablets USP 200mg Modafinil Tablets USP 200mg is a wakefulness-promoting agent used in the treatment of: Narcolepsy Obstructive Sleep Apnea (OSA) Shift Work Sleep Disorder (SWSD) Excessive Daytime Sleepiness (EDS) The formulation acts on specific neurotransmitters in the brain to enhance alertness and reduce extreme fatigue. Unlike traditional stimulants, Modafinil provides a smoother and more sustained wakefulness effect without excessive sedation or abrupt energy crashes. Mechanism of Action of Modafinil Tablets USP 200mg Modafinil works by influencing the central nervous […]
Cimetidine Tablets 400mg Manufacturer, Supplier & Exporter in India Introduction to Cimetidine Tablets 400mg Manufacturing Excellence We are a well-established manufacturer, exporter, and supplier of Cimetidine Tablets 400mg in India, delivering consistent pharmaceutical quality that meets international standards. Our commitment is to ensure high purity, precise formulation, and reliable therapeutic performance for global healthcare markets. Cimetidine Tablets 400mg are widely prescribed for the management of acid-related disorders, including gastroesophageal reflux disease (GERD), gastric ulcers, duodenal ulcers, and persistent heartburn. As a trusted pharmaceutical organization, we ensure that every batch is manufactured under strict quality control systems to guarantee safety, efficacy, and stability. Our manufacturing operations are based in Vapi, Gujarat, India, a well-recognized pharmaceutical hub known for advanced infrastructure and compliance-driven production facilities. Cimetidine Tablets 400mg Manufacturer in India Cimetidine Tablets We operate as a reputed pharmaceutical company focused on delivering high-quality generic formulations for domestic and international markets. Our strong manufacturing capability, research-driven processes, and regulatory compliance make us a preferred partner for global healthcare distributors. Taj Medico is recognized for its excellence in pharmaceutical manufacturing, supplying, and exporting a wide range of therapeutic medicines, including anti-ulcer, anti-infective, cardiovascular, and respiratory segments. Our core strengths include: WHO-GMP certified manufacturing […]
Glipizide 10mg Tablets Manufacturer Supplier in India – Trusted Exporter & PCD Pharma Franchise Company Leading Glipizide 10mg Tablets Manufacturer in India Taj Medico is a trusted name in the pharmaceutical industry, recognized as a leading manufacturer, exporter, and supplier of Glipizide 10 mg Tablets in India. We specialize in manufacturing high-quality antidiabetic medicines that meet international quality standards and regulatory requirements. Our Glipizide 10 mg Tablets are widely supplied across India as well as exported to Africa, the Middle East, LATAM countries, Southeast Asia, and various global markets. Glipizide 10 mg Tablets are prescribed for the treatment of type 2 diabetes mellitus and are used along with proper diet and exercise to control blood sugar levels effectively. At Taj Medico, we manufacture Glipizide Tablets under strict GMP-WHO manufacturing guidelines to ensure superior efficacy, safety, purity, and consistency. Glipizide 10 mg Tablets What is Glipizide 10mg Tablets? Glipizide belongs to the sulfonylurea class of antidiabetic medicines. It helps stimulate the pancreas to release more insulin, which lowers blood glucose levels naturally. Glipizide 10 mg Tablets are highly effective for adults suffering from type 2 diabetes and are commonly prescribed by healthcare professionals worldwide. Our premium quality Glipizide 10 mg Tablets […]
Darunavir Tablets 600mg Manufacturer Supplier in India Darunavir Ethanolate Tablets 600mg Manufacturer in India Taj Medico is recognized as a trusted pharmaceutical company engaged in the manufacturing, exporting, and supplying of high-quality Darunavir Ethanolate Tablets 600mg from Vapi, Gujarat, India. We specialize in manufacturing premium antiretroviral medicines that comply with international pharmaceutical standards and are widely supplied across Africa, the Middle East, LATAM, Asia, and other regulated and semi-regulated markets. Our commitment to pharmaceutical excellence, quality assurance, advanced manufacturing technology, and global distribution capabilities has positioned us among the leading exporters of Darunavir 600mg tablets in India. What is Darunavir Ethanolate Tablets 600mg? Darunavir Tablets 600mg Darunavir Ethanolate Tablets 600mg is an antiretroviral medication used for the treatment of HIV (Human Immunodeficiency Virus) infection. The medicine belongs to the class of drugs known as protease inhibitors, which work by blocking the protease enzyme required for viral replication. By reducing the amount of HIV in the body, Darunavir helps strengthen the immune system and decreases the risk of HIV-related complications and opportunistic infections. It is generally prescribed in combination with other antiretroviral medications, commonly including ritonavir, to improve therapeutic effectiveness. Mechanism of Action of Darunavir 600mg Tablets Darunavir acts by inhibiting […]
Darunavir Ethanolate Tablets 800mg Manufacturer, Supplier & Exporter in India Leading Manufacturer of Darunavir Ethanolate Tablets 800mg in India We are a reputed and globally recognized manufacturer, exporter, and supplier of Darunavir Ethanolate Tablets 800mg, delivering high-quality antiretroviral formulations from India to regulated and semi-regulated markets. With advanced pharmaceutical manufacturing infrastructure and strict compliance systems, we ensure consistent product quality, purity, and therapeutic effectiveness for global healthcare needs. Our manufacturing facility is designed to meet stringent international regulatory standards, enabling us to supply finished Darunavir Tablets to regions including Africa, Middle East, Latin America (LATAM), Southeast Asia, and CIS countries. We operate with a strong commitment to quality assurance, patient safety, and reliable supply chain management, making us a trusted partner in the HIV treatment segment. Darunavir Ethanolate Tablets 800mg – Therapeutic Overview Darunavir Tablets 800mg Darunavir Ethanolate Tablets 800mg belong to a class of medications known as HIV protease inhibitors. These medicines play a crucial role in antiretroviral therapy (ART) by blocking the protease enzyme required for viral replication. By inhibiting this enzyme, Darunavir helps: Reduce viral load in HIV-positive patients Improve immune system functioning Delay progression of HIV infection Reduce risk of HIV-related complications Improve life expectancy when […]
Sacubitril 49mg Valsartan 51mg Tablets Manufacturer in India Sacubitril 49mg Valsartan 51mg Tablets Manufacturer in India Taj Medico is recognized as a reliable and quality-focused pharmaceutical company engaged in the manufacturing, export, and supply of Sacubitril 49mg Valsartan 51mg Tablets in India. We specialize in delivering premium-quality cardiac medicines manufactured under strict international pharmaceutical standards for domestic and global healthcare markets. As a reputed pharmaceutical exporter from Vapi, Gujarat, India, we serve healthcare distributors, hospital suppliers, wholesalers, government agencies, pharmacies, and PCD franchise partners across Africa, the Middle East, LATAM, Asia, and CIS countries. Our advanced manufacturing capabilities and commitment to product quality make us one of the preferred exporters of Sacubitril & Valsartan Tablets worldwide. What Are Sacubitril 49mg Valsartan 51mg Tablets? Sacubitril Valsartan Tablets (Sacubvaltaj) Sacubitril 49mg Valsartan 51mg Tablets are widely prescribed for the treatment of chronic heart failure with reduced ejection fraction (HFrEF). This combination medicine helps improve heart function, reduce blood pressure, decrease hospitalization risk, and lower cardiovascular mortality in heart failure patients. The medicine combines two active ingredients: Sacubitril 49mg – Helps relax blood vessels and improve sodium excretion. Valsartan 51mg – An angiotensin receptor blocker (ARB) that lowers blood pressure and reduces stress […]
Sacubitril & Valsartan Tablets 50mg Manufacturer in India Sacubitril & Valsartan Tablets 50mg Manufacturer and Supplier in India Taj Medico is recognized as a reliable and professional pharmaceutical company engaged in the manufacturing, supply, export, and distribution of Sacubitril & Valsartan Tablets 50mg across domestic and international markets. We manufacture premium-quality cardiac medicines under stringent pharmaceutical standards to meet the growing global demand for effective heart failure treatment medications. Our advanced manufacturing facilities in Vapi are equipped with modern technology, highly controlled production systems, and strict quality assurance procedures. We supply Sacubitril & Valsartan Tablets 50mg to hospitals, government institutions, healthcare distributors, pharmacies, wholesalers, importers, and PCD pharma franchise companies across India, Africa, the Middle East, LATAM countries, Southeast Asia, and other regulated and semi-regulated markets. What is Sacubitril & Valsartan Tablets 50mg? Sacubitril Valsartan Tablets 50mg Sacubitril & Valsartan Tablets 50mg is a combination medicine widely prescribed for the treatment of chronic heart failure in adults. The medicine combines two active pharmaceutical ingredients: Sacubitril Valsartan This combination helps improve cardiac efficiency, reduce strain on the heart, decrease hospitalization risk, and improve survival rates in patients suffering from chronic heart failure with reduced ejection fraction (HFrEF). We manufacture this formulation […]
Captopril 12.5mg Tablets Manufacturer Supplier in India Leading Captopril 12.5mg Tablets Manufacturer in India Taj Medico is recognized as a leading pharmaceutical company engaged in the manufacturing, supplying, and exporting of high-quality Captopril 12.5mg Tablets under the brand name Capotaj. We are committed to delivering pharmaceutical formulations that meet international quality standards and fulfill the growing demand for cardiovascular medicines across domestic and global markets. Our Captopril 12.5mg Tablets are manufactured in advanced WHO-GMP compliant facilities using premium-grade raw materials and stringent quality control procedures. We supply our products across PAN India and export to multiple countries in Africa, the Middle East, LATAM, Asia, and other international pharmaceutical markets. What is Captopril 12.5mg Tablet? Captopril 12.5mg Tablet Captopril 12.5mg Tablets belong to the category of medicines known as ACE inhibitors (Angiotensin-Converting Enzyme inhibitors). This medication is commonly prescribed for the treatment of: High blood pressure (Hypertension) Congestive heart failure Diabetic kidney disease Prevention of heart attack and stroke Improving survival after myocardial infarction Captopril works by relaxing blood vessels, improving blood circulation, and reducing strain on the heart. It is one of the most trusted medications used worldwide for cardiovascular care. Why Choose Taj Medico for Captopril 12.5mg Tablets? WHO-GMP […]
Captopril 50mg Tablets Manufacturer and Supplier in India – Trusted Exporter from Vapi, Gujarat Leading Captopril 50mg Tablets Manufacturer in India Taj Medico is recognized as a reliable and quality-driven pharmaceutical company engaged in the manufacturing, supplying, and exporting of Captopril 50mg Tablets from Vapi, Gujarat, India. We are committed to delivering premium-quality cardiovascular medicines that comply with international pharmaceutical standards for safety, efficacy, and purity. As a reputed manufacturer and exporter of Captopril Tablets, we cater to domestic and international pharmaceutical markets including Africa, the Middle East, LATAM countries, Southeast Asia, and CIS nations. Our advanced manufacturing facilities and strict quality assurance systems help us maintain consistency in every batch. What is Captopril 50mg Tablet? Captopril is an angiotensin-converting enzyme (ACE) inhibitor widely prescribed for the treatment of high blood pressure and heart-related conditions. It works by relaxing blood vessels and improving blood circulation, allowing the heart to pump blood more efficiently. Captopril Tablets are commonly recommended for: Hypertension (High Blood Pressure) Congestive Heart Failure Post Heart Attack Recovery Diabetic Kidney Disease Prevention of Stroke and Cardiovascular Complications The medicine helps lower blood pressure effectively and reduces the risk of heart attack, kidney damage, and other cardiovascular disorders. Why […]
Captopril 25mg Tablets Manufacturer and Supplier in India Captopril 25mg Tablets Manufacturer in India Taj medico is recognized as a leading pharmaceutical company specializing in the manufacturing, supply, and export of high-quality Captopril 25mg Tablets under the brand name Capotaj. Based in Vapi, we have established a strong reputation for delivering reliable cardiovascular medicines that comply with international quality standards and regulatory requirements. Our Captopril 25mg Tablets are widely supplied across India and exported to countries in Africa, the Middle East, Latin America, and Asian markets. We focus on delivering premium-quality formulations with consistent therapeutic effectiveness, superior packaging standards, and timely global distribution support. What is Captopril 25mg Tablet? Captopril is an ACE inhibitor medicine used for the treatment of hypertension, congestive heart failure, diabetic nephropathy, and left ventricular dysfunction after myocardial infarction. It works by relaxing blood vessels, improving blood flow, and reducing the workload on the heart. Captopril 25mg Tablets are prescribed by healthcare professionals to help patients maintain healthy blood pressure levels and reduce the risk of cardiovascular complications. Composition of Capotaj Captopril Tablets Product NameCapotajGeneric NameCaptopril TabletsStrength25mgDosage FormTabletsPackagingBlister / Alu-AluTherapeutic CategoryAnti-HypertensiveManufactured ByTaj medicoOriginIndia Benefits of Captopril 25mg Tablets Captopril Tablets 25mg (CAPOTAJ) Effective Blood Pressure Control […]
Liothyronine 20mcg Tablets Manufacturer Supplier in India Liothyronine 20mcg Tablets Manufacturer in India Taj medico is recognized as a trusted pharmaceutical manufacturing company specializing in premium-quality Liothyronine 20mcg Tablets under the brand name LUKOI-T3. We are committed to manufacturing high-purity thyroid hormone formulations that meet international pharmaceutical standards for safety, efficacy, and consistency. As a reputed pharmaceutical company based in Vapi, we supply and export Liothyronine Tablets across India, Africa, the Middle East, LATAM countries, Southeast Asia, and global regulated and semi-regulated markets. Our advanced manufacturing infrastructure, stringent quality systems, and customer-focused approach have established us among the leading Liothyronine 20mcg Tablets manufacturers and exporters from India. What is Liothyronine 20mcg Tablet? Liothyronine 20mcg Tablets Liothyronine 20mcg Tablets contain Liothyronine Sodium, a synthetic form of the thyroid hormone T3 (Triiodothyronine). It is prescribed primarily for the treatment of hypothyroidism, a condition in which the thyroid gland fails to produce sufficient thyroid hormones required for maintaining metabolism and overall body function. Liothyronine acts rapidly and helps regulate: Metabolic rate Body temperature Energy production Heart function Muscle control Brain development Digestive function Our LUKOI-T3 tablets are manufactured using pharmaceutical-grade active ingredients and comply with strict quality control measures to ensure accurate potency […]
Warfarin Sodium 3mg Tablets Manufacturer Supplier in India Warfarin Sodium 3mg Tablets Manufacturer in India Taj medico is recognized as a leading pharmaceutical company engaged in the manufacturing, supplying, and exporting of high-quality Warfarin Sodium 3mg Tablets from India. We manufacture pharmaceutical formulations under strict international quality standards to ensure safety, efficacy, purity, and consistent therapeutic performance for global healthcare markets. Warfarin Sodium 3mg Tablets are widely prescribed anticoagulant medicines used for preventing and treating blood clot-related disorders. Our formulation is manufactured using premium-grade active pharmaceutical ingredients and advanced production technologies in WHO-GMP compliant facilities. We serve hospitals, distributors, government tenders, pharmacies, institutional buyers, and healthcare organizations across Africa, the Middle East, LATAM, Asia, and other international markets. Our commitment to pharmaceutical excellence, timely delivery, competitive pricing, and export-quality packaging has established us as a reliable Warfarin Sodium 3mg Tablets manufacturer and exporter in India. What is Warfarin Sodium 3mg Tablets? Warfarin Sodium 3mg (Rifataj) Warfarin Sodium 3mg Tablets are oral anticoagulant medicines that help prevent the formation of harmful blood clots inside blood vessels. The medicine works by reducing the clotting ability of blood and lowering the risk of severe cardiovascular complications. Warfarin is commonly prescribed for patients suffering […]
Oxycodone Hydrochloride 30mg Tablets Manufacturer, Supplier & Exporter in India Trusted Oxycodone Hydrochloride 30mg Tablets Manufacturing Excellence in India We are a reputed and quality-driven manufacturer, exporter, and supplier of Oxycodone Hydrochloride 30mg Tablets in India, delivering globally compliant pharmaceutical solutions for pain management therapies. Our manufacturing facility is based in Vapi, Gujarat, and operates under strict quality-controlled environments designed to meet international regulatory expectations. We focus on delivering consistent potency, high purity, and stable formulation standards that ensure safe therapeutic performance under medical supervision. Our Oxycodone Hydrochloride 30mg tablets are developed using advanced formulation technology and tested across multiple quality checkpoints before final dispatch. About Oxycodone Hydrochloride 30mg Tablets Oxycodone Hydrochloride 30mg Tablets Oxycodone Hydrochloride 30mg Tablets belong to the class of opioid analgesics used in the management of moderate to severe pain conditions where alternative pain relief therapies are insufficient. These tablets act on the central nervous system (CNS) by binding to opioid receptors, helping reduce the perception of pain and improving patient comfort under prescribed clinical conditions. Due to its strong analgesic profile, Oxycodone is used strictly under medical supervision in controlled dosage forms. We ensure that every batch is manufactured with precise pharmaceutical standards to support […]
Oxycodone Hydrochloride 20mg Tablets Manufacturer Supplier in India Leading Oxycodone Hydrochloride 20mg Tablets Manufacturer in India Taj Pharma is recognized among the leading pharmaceutical manufacturing companies engaged in the production, supply, and export of high-quality Oxycodone Hydrochloride 20mg Tablets from India. We manufacture premium-grade opioid analgesic medicines under strict pharmaceutical quality standards for regulated and semi-regulated international markets. As a trusted pharmaceutical exporter from Vapi, we supply Oxycodone Hydrochloride 20mg Tablets to healthcare distributors, hospital suppliers, government agencies, pharmacies, and licensed pharmaceutical importers across Africa, the Middle East, LATAM countries, Southeast Asia, and global markets. Our commitment to pharmaceutical excellence, consistent product quality, regulatory compliance, and timely international delivery has positioned us as a reliable manufacturer and exporter of opioid pain management medicines from India. What is Oxycodone Hydrochloride 20mg Tablets? Oxycodone Hydrochloride 20mg Tablets Oxycodone Hydrochloride 20mg Tablets is a potent opioid analgesic medicine used for the management of moderate to severe pain where alternative treatments are ineffective or not tolerated. Oxycodone works directly on the central nervous system to relieve pain and improve patient comfort in chronic and acute pain conditions. The medicine is commonly prescribed for: Cancer-related pain Post-surgical pain Severe musculoskeletal pain Trauma-related pain Neuropathic pain […]
Oxycodone Hydrochloride 80mg Tablets Manufacturer Supplier in India Leading Oxycodone Hydrochloride 80mg Tablets Manufacturer in India Taj Medico is recognized among the leading pharmaceutical manufacturing companies for premium-quality Oxycodone Hydrochloride 80mg Tablets manufacturing, supply, and global export services. We specialize in the production of high-strength opioid analgesic formulations designed for effective management of severe chronic pain in opioid-tolerant patients. Our manufacturing operations in Vapi, Gujarat, India are equipped with advanced pharmaceutical technology, stringent quality assurance systems, and internationally compliant production standards. We deliver reliable pharmaceutical solutions to regulated and semi-regulated markets across Africa, the Middle East, LATAM, Southeast Asia, and other global regions. What Are Oxycodone Hydrochloride 80mg Tablets? Oxycodone Hydrochloride 80mg Tablets Oxycodone Hydrochloride 80mg Tablets are high-potency opioid analgesics prescribed for patients experiencing severe pain requiring continuous, around-the-clock opioid treatment over an extended period. These tablets are commonly used when alternative pain management therapies fail to provide adequate relief. The medication works by altering the brain and nervous system’s response to pain. Extended-release Oxycodone formulations are specifically developed to provide long-lasting pain control in chronic conditions requiring sustained analgesic support. Medical Uses of Oxycodone Hydrochloride 80mg Tablets Oxycodone Hydrochloride 80mg Tablets are widely prescribed for: Severe chronic pain […]
Oxycodone Hydrochloride 40mg Tablets Manufacturer and Exporter in India Trusted Oxycodone Hydrochloride 40mg Tablets Manufacturer in India Taj Medico is recognized as a leading pharmaceutical company engaged in the manufacturing, supplying, and exporting of high-quality Oxycodone Hydrochloride 40mg Tablets from Vapi, Gujarat, India. We are committed to maintaining strict pharmaceutical quality standards, ensuring purity, safety, efficacy, and global compliance in every batch we manufacture. Our advanced manufacturing facilities, experienced pharmaceutical professionals, and international-quality packaging standards make us one of the most reliable Oxycodone Tablets manufacturers and exporters in India. We cater to pharmaceutical importers, hospitals, distributors, government tenders, and healthcare institutions across Africa, the Middle East, LATAM, Asia, and regulated international markets. What Are Oxycodone Hydrochloride 40mg Tablets? Oxycodone Hydrochloride 40mg Tablets Oxycodone Hydrochloride 40mg Tablets are powerful prescription opioid analgesics used for the management of moderate to severe chronic pain requiring long-term, around-the-clock treatment. The medicine belongs to the opioid pain medication class and works directly on the central nervous system to alter the body’s response to pain. Oxycodone hydrochloride is synthesized by modifying thebaine, a naturally occurring compound extracted from the opium poppy plant. It binds to mu-opioid receptors in the brain and spinal cord, effectively blocking pain […]
Warfarin Sodium 1mg Tablets Manufacturer Supplier in India Warfarin Sodium 1mg Tablets Manufacturer in India Taj Medico is a trusted name in the pharmaceutical industry, recognized as a leading manufacturer, supplier, and exporter of high-quality Warfarin Sodium 1mg Tablets from India. We manufacture pharmaceutical formulations under strict quality standards to ensure safety, efficacy, purity, and international compliance for regulated and semi-regulated markets. Warfarin Sodium 1mg Tablets are widely prescribed anticoagulant medicines used to prevent and treat harmful blood clots. These tablets are essential in the management of conditions such as deep vein thrombosis (DVT), pulmonary embolism, atrial fibrillation, and stroke prevention. As a reputed pharmaceutical exporter from Vapi, Gujarat, India, we supply premium-quality Warfarin tablets to buyers across Africa, the Middle East, LATAM countries, Southeast Asia, and global pharmaceutical markets. What is Warfarin Sodium 1mg Tablet? Warfarin Sodium 1mg Tablets Warfarin Sodium 1mg Tablet is an oral anticoagulant medicine commonly known as a blood thinner. It works by reducing the ability of blood to clot, thereby preventing dangerous clot formation in veins, arteries, lungs, brain, and heart. Doctors prescribe Warfarin tablets for patients suffering from: Deep vein thrombosis (DVT) Pulmonary embolism Atrial fibrillation Stroke prevention Artificial heart valve complications Blood […]
Fenbendazole Tablets 222mg Manufacturer in India – Leading Supplier, Exporter & PCD Pharma Franchise Company In today’s rapidly growing veterinary pharmaceutical market, finding a trusted Fenbendazole Tablets 222mg Manufacturer in India is essential for distributors, importers, wholesalers, veterinary hospitals, and pharma franchise companies. At Taj Medico, we proudly position ourselves as a leading manufacturer, exporter, supplier, and PCD pharma franchise provider of premium-quality Fenbendazole Tablets 222mg (Orray) from Vapi. We manufacture high-quality Fenbendazole Tablets 222mg using advanced pharmaceutical technology, premium-grade raw materials, and strict quality control systems to ensure superior effectiveness, purity, and safety. Our products are supplied across PAN India and exported to major international markets including Africa, the Middle East, LATAM, Asia, and Europe. ✔ Premium Quality Manufacturing✔ Bulk Supply Availability✔ Global Export Support✔ PCD Pharma Franchise Opportunities✔ Third Party Manufacturing Services✔ Affordable Pricing Structure 💊 What is Fenbendazole Tablets 222mg (Orray)? Fenbendazole Tablets Fenbendazole is a broad-spectrum antiparasitic medication widely used to treat parasitic infections caused by worms and intestinal parasites. It belongs to the benzimidazole group of medicines and is commonly used in veterinary healthcare. Fenbendazole Tablets 222mg are commonly used for: ✔ Roundworms✔ Hookworms✔ Tapeworms✔ Lungworms✔ Whipworms✔ Giardia infections✔ Intestinal parasitic infections Our Fenbendazole Tablets […]